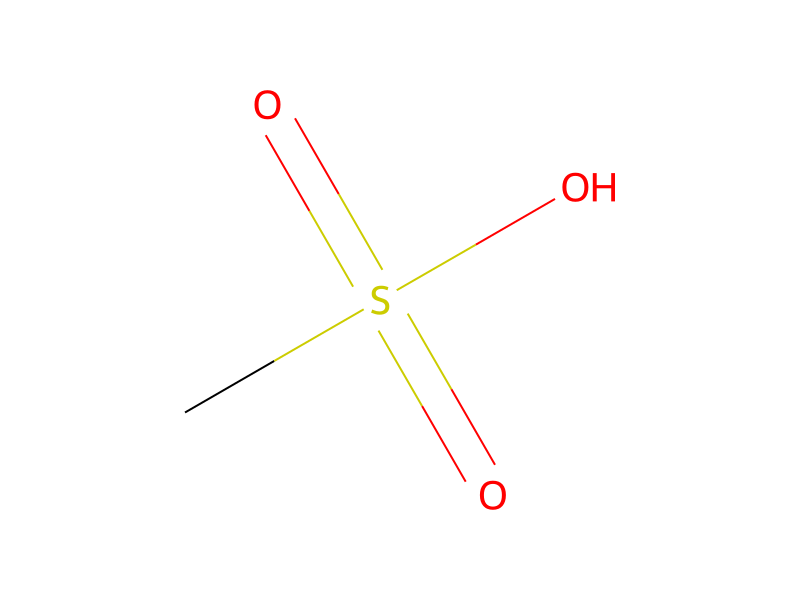
Methanesulfonic Acid → HCID 178493015, compound
Reaction equation

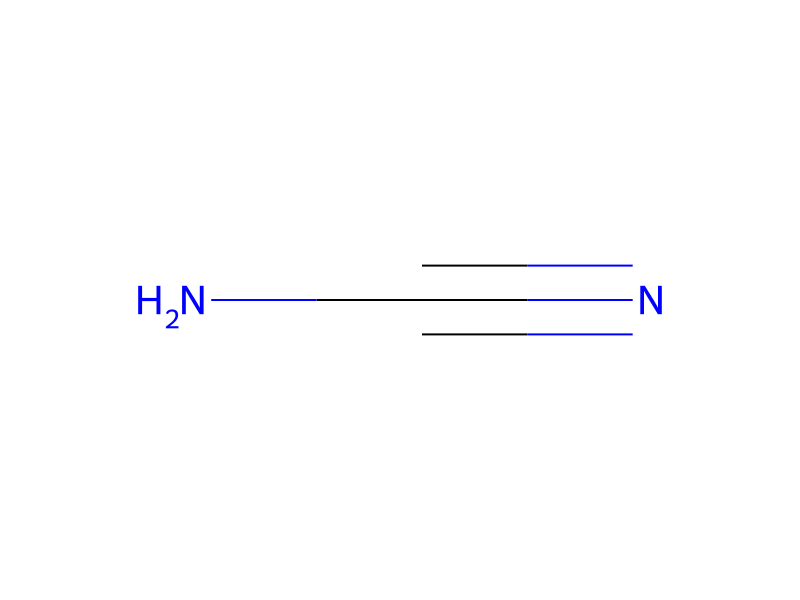
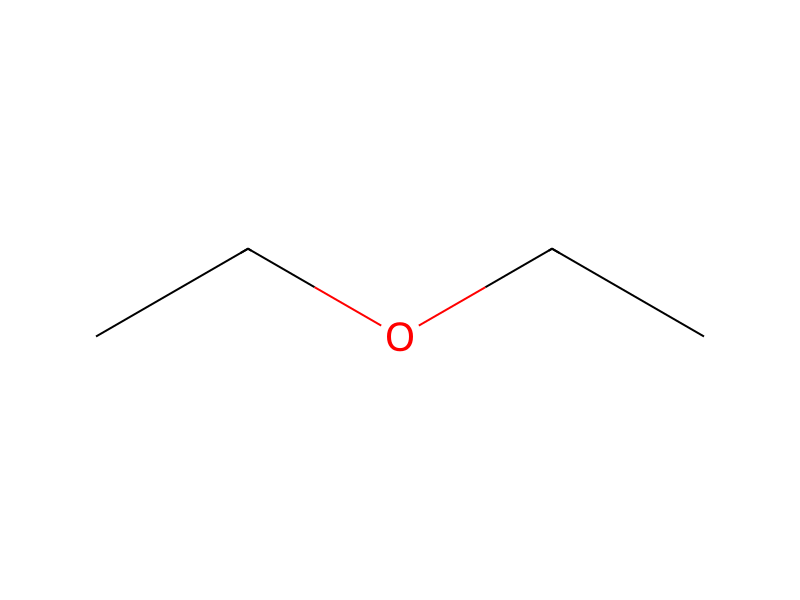
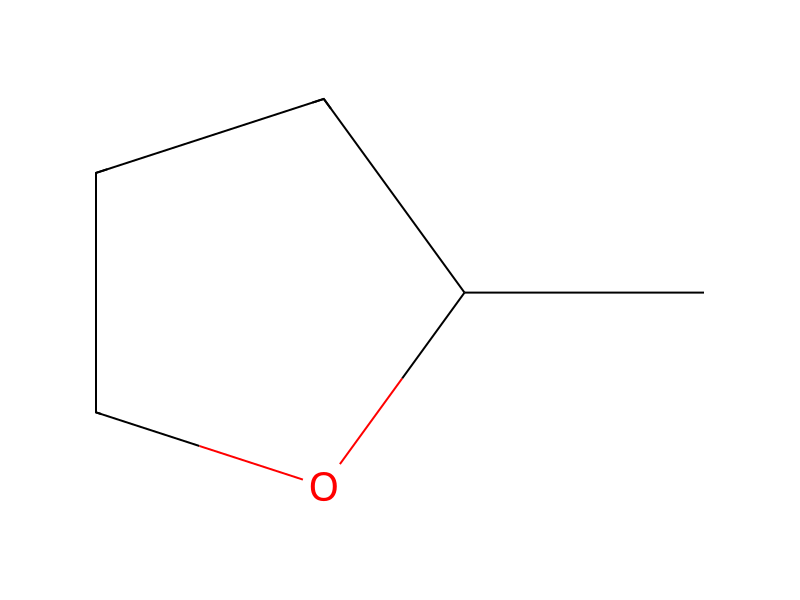
CS(O)(=O)=O.[CH3:6][O:7][C:8]1[CH:14]=[C:13]([N:15]2[CH2:20][CH2:19][N:18]([CH3:21])[CH2:17][CH2:16]2)[C:12]([N+:22]([O-:24])=[O:23])=[CH:11][C:9]=1[NH2:10].[N:25]#[C:26][NH2:27].[OH-].[Na+]>C(O)CCC.O.CC1CCCO1.C(OCC)C>[CH3:6][O:7][C:8]1[CH:14]=[C:13]([N:15]2[CH2:20][CH2:19][N:18]([CH3:21])[CH2:17][CH2:16]2)[C:12]([N+:22]([O-:24])=[O:23])=[CH:11][C:9]=1[NH:10][C:26]([NH2:27])=[NH:25]Participants
Reactant 4

Product 1

Reagents, catalysts & solvents 4

Conditions
- Temperature
- 90 °C
Procedure
Methanesulfonic acid (0.508 mL, 7.83 mmol) was added to a slurry of 2-methoxy-4-(4-methylpiperazin-1-yl)-5-nitroaniline (Intermediate 14, 1.39 g, 5.22 mmol) in butan-1-ol (10 mL) and water (0.5 mL) at r.t. The resulting slurry was heated to 90° C. and at that temperature a solution of cyanamide (0.439 g, 10.44 mmol) in water (0.22 mL) was added dropwise over a period of 1 minute. The mixture was heated at 90° C. for 0.5 h then methanesulfonic acid (0.339 mL, 5.22 mmol) was added dropwise. At 10-minute intervals more cyanamide (0.219 g, 5.22 mmol), more methanesulfonic acid (0.339 mL, 5.22 mmol) and more cyanamide (0.329 g, 7.83 mmol) and more methanesulfonic acid (0.339 mL, 5.22 mmol) were successively added. The mixture was allowed to cool and was diluted with 2-methyltetrahydrofuran (75 mL) and diethyl ether (75 mL). This solution was then basified using 5M NaOH to ˜pH 13. The resulting phases were separated and the aqueous solution was extracted with 2-methyltetrahydrofuran (3×50 mL). The combined organic solutions were washed with sat. brine and dried (MgSO4). The product precipitated onto the drying agent, so after filtration the MgSO4 was washed with hot CH2Cl2/CH3OH 5:1 (5×100 mL) and the combined filtrate was concentrated in vacuo. The residue was dissolved in hot 2-s propanol (80 mL), filtered while hot and then concentrated in vacuo. The residue was dissolved in CH2Cl2 (50 mL), filtered and the filtrate was concentrated in vacuo to give the title compound (1.49 g, 89%) as an orange foam. The product could only be deposited amorphously from the 2-propanol filtrate, but a 38 mg sample of the final product was crystallised from ethanol (˜100 μL) and was collected by filtration, washed with −70° C. ethanol and diethyl ether and dried under vacuum to give the title compound (11 mg, 28%) as a yellow powder; 1H NMR: 2.25 (3H, s), 2.44-2.49 (4H, m), 2.99-3.16 (4H, m), 3.91 (3H, s), 6.73 (1H, s), 7.38 (3H, s), 7.67 (1H, s); m/z: ES+ MH+ 309.
Workup
- additionwas added dropwise over a period of 1 minute
- temperatureto cool
- customThe resulting phases were separated
- extractionthe aqueous solution was extracted with 2-methyltetrahydrofuran (3×50 mL)
- washThe combined organic solutions were washed with sat. brine
- dry with materialdried (MgSO4)
- customThe product precipitated onto the drying agent
- filtrationso after filtration the MgSO4
- washwas washed with hot CH2Cl2/CH3OH 5:1 (5×100 mL)
- concentrationthe combined filtrate was concentrated in vacuo
- dissolutionThe residue was dissolved in hot 2-s propanol (80 mL)
- filtrationfiltered while hot and
- concentrationthen concentrated in vacuo
- dissolutionThe residue was dissolved in CH2Cl2 (50 mL)
- filtrationfiltered
- concentrationthe filtrate was concentrated in vacuo
Sources & evidence
- ORD record
- ord-1c38296e7f314f2faa11475d6ce83230
- Source dataset
- uspto-grants-2015_02
- Patent number
- US08946235B2
Reaction notes
Loading…