Dichloromethane → HCID 53305815, compound
Reaction equation

ClCCl.C1(P(C2C=CC=CC=2)C2C=CC=CC=2)C=CC=CC=1.[C:23]([O:31][CH2:32][C@@H:33]1[C@@H:37]([O:38][C:39](=[O:46])[C:40]2[CH:45]=[CH:44][CH:43]=[CH:42][CH:41]=2)[C@:36]([F:48])([CH3:47])[C@H:35](O)[O:34]1)(=[O:30])[C:24]1[CH:29]=[CH:28][CH:27]=[CH:26][CH:25]=1.C(Br)(Br)(Br)[Br:51]>CCOC(C)=O>[C:23]([O:31][CH2:32][C@@H:33]1[C@@H:37]([O:38][C:39](=[O:46])[C:40]2[CH:45]=[CH:44][CH:43]=[CH:42][CH:41]=2)[C@:36]([F:48])([CH3:47])[C@@H:35]([Br:51])[O:34]1)(=[O:30])[C:24]1[CH:29]=[CH:28][CH:27]=[CH:26][CH:25]=1Participants
Reactant 4


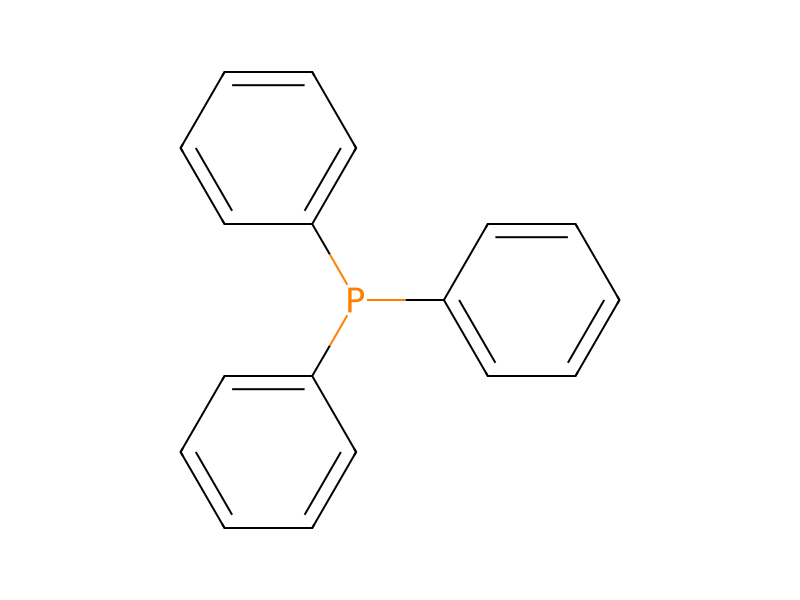

Product 1

Reagents, catalysts & solvents 1
Conditions
- Temperature
- -17 °C
Procedure
Anhydrous dichloromethane (5.6 L) was charged into a reactor and cooled to −22° C. or below. Triphenylphosphine (205.4 g, 0.783 mol) was added to the cold solvent and the suspension was stirred to form a solution. The lactol (2, 209.4 g, 0.559 mol) in solid form was added to the cold solution and stirred for 15 mins. Carbon tetrabromide (278.2 g, 0.839 mol) was added portion-wise while maintaining the temperature of the solution between −22° C. to −20° C. under a flow of nitrogen gas (approx. 30 min). After finishing the addition of CBr4, the temperature was slowly raised to −17° C. over 20 mins. The reaction was judged to be >95% complete by TLC (Rfs 0.61 (α), 0.72 ((β), 0.36 lactol; 20% EtOAc in hexanes). The reaction solution was immediately transferred to a vessel containing 230 g of flash chromatography grade silica gel (40-63 microns). The stirred mixture was immediately passed through a pad of silica gel (680 g) in a 2.5 L sintered glass Buchner funnel. The filtrate was concentrated under reduced pressure to about 800 mL and the ratio of α/β isomers of the crude product was 10:1 as determined by 1H-NMR. (CDCl3) δ=6.35, (s, α C1-H), 6.43, (d, β C1-H). The residue was purified by plug column chromatography using 2.1 kg of silica gel in a 6 L sintered glass Buchner funnel and eluted (via suction) with a stepwise gradient elution of 1%, 5%, 8% 12% EtOAc in hexane (ca 4 L each) to remove non-polar impurities followed by 12%, 25% EtOAc in hexane (6 L total) to elute the product. The product containing fractions were combined into two fractions, concentrated under reduced pressure, dried under vacuum (0.1 mmHg, ambient temp., 20 h) to colorless oils. Main fraction (197 g, 89% α/β=20:1). The alpha isomer crystallized from a small portion of the oil upon standing at 0° C. for several weeks to give large, thin plates, mp 59-61° C. The pure beta isomer crystallized from a mixture of alpha and beta product oil from an earlier less selective run to give needles, mp 77-79° C.
Workup
- temperaturecooled to −22° C. or below
- customto form a solution
- stirringstirred for 15 mins
- temperaturewhile maintaining the temperature of the solution between −22° C. to −20° C. under a flow of nitrogen gas (approx. 30 min)
- customThe reaction solution was immediately transferred to a vessel
- additioncontaining 230 g of flash chromatography grade silica gel (40-63 microns)
- concentrationThe filtrate was concentrated under reduced pressure to about 800 mL
- customThe residue was purified by plug column chromatography
- washeluted (via suction)
- washwith a stepwise gradient elution of 1%, 5%, 8% 12% EtOAc in hexane (ca 4 L each)
- customto remove non-polar impurities
- washto elute the product
- additionThe product containing fractions
- concentrationconcentrated under reduced pressure
- customdried under vacuum (0.1 mmHg, ambient temp., 20 h) to colorless oils
- customThe alpha isomer crystallized from a small portion of the oil
- waitupon standing at 0° C. for several weeks
- customto give large
- customThe pure beta isomer crystallized from a mixture of alpha and beta product oil from an earlier less selective run
- customto give needles, mp 77-79° C.
Sources & evidence
- ORD record
- ord-d0d2994db94048bda5e09c910f1a389b
- Source dataset
- uspto-grants-2015_02
- Patent number
- US08957045B2
Reaction notes
Loading…