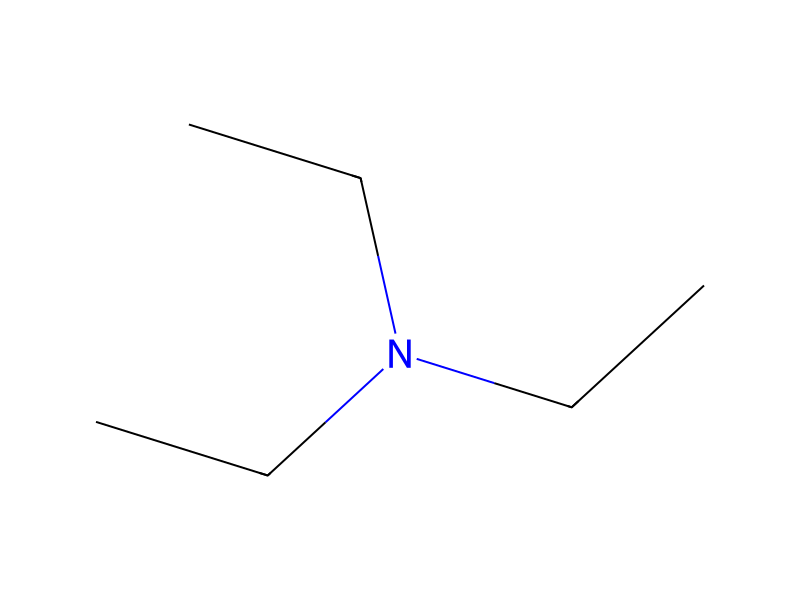
Triethylamine → HCID 67491161, compound
Reaction equation

C(N(CC)CC)C.F[C:9]1[C:18]([CH3:19])=[C:17]2[C:12]([C:13](=[O:27])[C:14]([C:24]([OH:26])=[O:25])=[CH:15][N:16]2[C@@H:20]2[CH2:22][C@@H:21]2[F:23])=[CH:11][CH:10]=1.C(OC([NH:35][C@:36]12[CH2:44][NH:43][CH2:42][C@@H:41]1[CH2:40][O:39][CH2:38][CH2:37]2)=O)(C)(C)C>CS(C)=O.C(OCC)(=O)C>[NH2:35][C@:36]12[CH2:44][N:43]([C:9]3[C:18]([CH3:19])=[C:17]4[C:12]([C:13](=[O:27])[C:14]([C:24]([OH:26])=[O:25])=[CH:15][N:16]4[C@@H:20]4[CH2:22][C@@H:21]4[F:23])=[CH:11][CH:10]=3)[CH2:42][C@@H:41]1[CH2:40][O:39][CH2:38][CH2:37]2Participants
Reactant 3


Product 1

Reagents, catalysts & solvents 2
Conditions
- Temperature
- 75 °C
Procedure
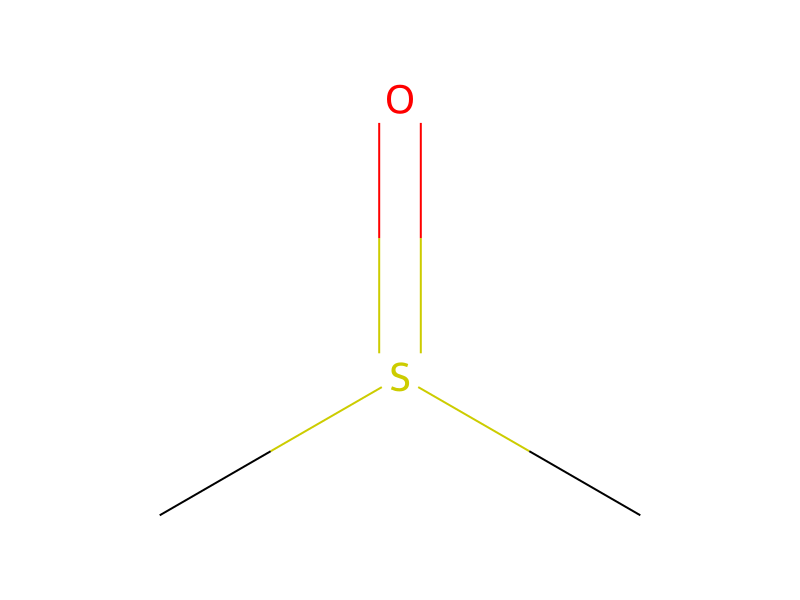
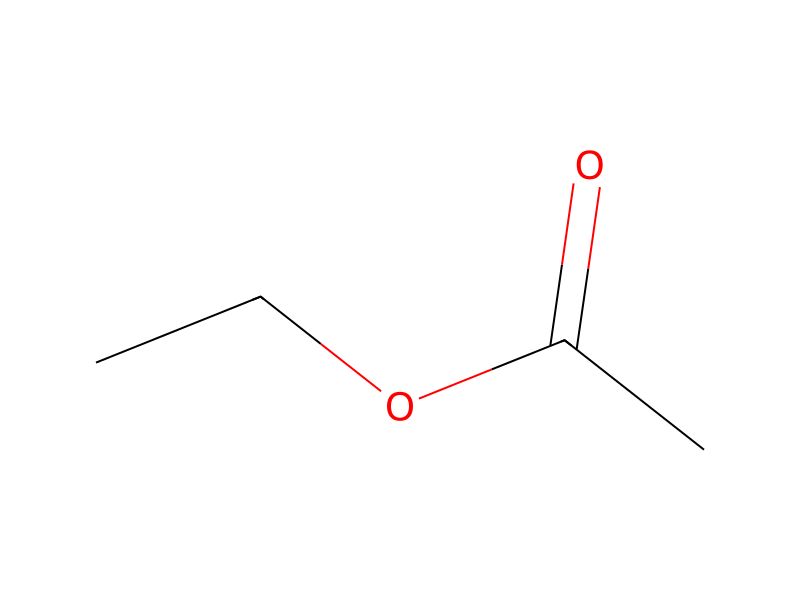
Triethylamine (0.0737 ml, 0.528 mmol) and 7-fluoro-1-[(1R,2S)-2-fluorocyclopropyl]-1,4-dihydro-8-methyl-4-oxoquinoline-3-carboxylic acid (49.0 mg, 0.175 mmol) were added to a (1S,6S)-1-tert-butoxycarbonylamino-4-oxa-8-azabicyclo[4.3.0]nonane (42.2 mg, 0.176 mmol) in dimethyl sulfoxide (0.352 mL) in a nitrogen atmosphere, and the mixture was stirred at 75° C. for two days. Dimethyl sulfoxide (0.352 mL) and triethylamine (0.147 mL, 1.06 mmol) were added to the reaction solution, and the mixture was stirred at 75° C. for four days. Dimethyl sulfoxide (0.352 mL) and triethylamine (0.147 ml, 1.06 mmol) were added to the reaction solution, and the mixture was stirred at 75° C. for two days. 7-Fluoro-1-[(1R,2S)-2-fluorocyclopropyl]-1,4-dihydro-8-methyl-4-oxoquinoline-3-carboxylic acid (24.6 mg, 0.0881 mmol) and triethylamine (0.147 mL, 1.06 mmol) were added to the reaction solution, and the mixture was stirred for five days. The reaction solution was diluted with ethyl acetate (30 mL) and then washed with a 10% citric acid solution (25 mL), water (25 mL), and a saturated sodium hydroxide solution (25 mL). Further, the organic layer was extracted again from the washing 10% citric acid solution and the washing water with ethyl acetate (40 mL). The organic layers were combined, dried over anhydrous sodium sulfate, and filtered, and then the filtrate was concentrated under reduced pressure. The resulting residue was purified by silica gel column chromatography (chloroform:methanol=100:0→99:1→98:2). Trifluoroacetic acid (0.402 mL, 3 v/w) was added to a solution of 67.5 mg of the resulting residue in dichloromethane (1.32 mL) with stirring under ice-cooling, and the mixture was stirred at room temperature for one hour. The reaction solution was concentrated under reduced pressure, and trifluoroacetic acid was azeotropically distilled with toluene (×3). 6 mol/L hydrochloric acid was added to the resulting residue under ice-cooling, and the mixture was washed with chloroform (30 mL×5). The aqueous layer was adjusted to pH 12 with a 1 mol/L sodium hydroxide solution under ice-cooling. The basic solution was adjusted to pH 7.4 with 1 mol/L hydrochloric acid, followed by extraction with chloroform (100 mL×2) and chloroform/methanol=10/1 (50 mL). The organic layer was dried over anhydrous sodium sulfate and filtered, and then the filtrate was concentrated under reduced pressure. The residue was purified by PTLC (lower layer of chloroform/methanol/water=7/3/1) and concentrated under reduced pressure. The resulting residue was dissolved in hot ethanol, and the insoluble material was removed by filtration. The filtrate was concentrated under reduced pressure, dissolved in hot ethanol (1 mL), ultrasonically treated, and cooled with ice water. Then, the slurry was washed with diethyl ether (10 mL). After stirring at room temperature overnight, the precipitated crystals were collected by filtration, washed with ethanol and diethyl ether, and then dried under reduced pressure at 60° C. to give 12.4 mg of the title compound.
Workup
- stirringthe mixture was stirred at 75° C. for four days
- stirringthe mixture was stirred at 75° C. for two days
- stirringthe mixture was stirred for five days
- washwashed with a 10% citric acid solution (25 mL), water (25 mL)
- extractionFurther, the organic layer was extracted again from the washing 10% citric acid solution
- dry with materialdried over anhydrous sodium sulfate
- filtrationfiltered
- concentrationthe filtrate was concentrated under reduced pressure
- customThe resulting residue was purified by silica gel column chromatography (chloroform:methanol=100:0→99:1→98:2)
- additionTrifluoroacetic acid (0.402 mL, 3 v/w) was added to a solution of 67.5 mg of the resulting residue in dichloromethane (1.32 mL)
- stirringwith stirring under ice-
- temperaturecooling
- stirringthe mixture was stirred at room temperature for one hour
- concentrationThe reaction solution was concentrated under reduced pressure, and trifluoroacetic acid
- distillationwas azeotropically distilled with toluene (×3)
- addition6 mol/L hydrochloric acid was added to the resulting residue under ice-
- temperaturecooling
- washthe mixture was washed with chloroform (30 mL×5)
- temperaturecooling
- extractionfollowed by extraction with chloroform (100 mL×2) and chloroform/methanol=10/1 (50 mL)
- dry with materialThe organic layer was dried over anhydrous sodium sulfate
- filtrationfiltered
- concentrationthe filtrate was concentrated under reduced pressure
- customThe residue was purified by PTLC (lower layer of chloroform/methanol/water=7/3/1)
- concentrationconcentrated under reduced pressure
- dissolutionThe resulting residue was dissolved in hot ethanol
- customthe insoluble material was removed by filtration
- concentrationThe filtrate was concentrated under reduced pressure
- dissolutiondissolved in hot ethanol (1 mL)
- additionultrasonically treated
- temperaturecooled with ice water
- washThen, the slurry was washed with diethyl ether (10 mL)
- stirringAfter stirring at room temperature overnight
- filtrationthe precipitated crystals were collected by filtration
- washwashed with ethanol and diethyl ether
- customdried under reduced pressure at 60° C.
Sources & evidence
- ORD record
- ord-f1ecb45d5b5a4c41ae16b3aaf6310423
- Source dataset
- uspto-grants-2013_12
- Patent number
- US08618094B2
Reaction notes
Loading…