Ethyl Acetate → HCID 58438087, compound
Reaction equation

Br[CH:2]([CH2:7][CH2:8][CH2:9]Br)[C:3]([O:5][CH3:6])=[O:4].C([O-])([O-])=O.[K+].[K+].[Br:17][C:18]1[CH:23]=[CH:22][C:21]([C@@H:24]([NH2:26])[CH3:25])=[CH:20][CH:19]=1.CCOC(C)=O>CC#N.O>[CH3:6][O:5][C:3]([C@@H:2]1[CH2:7][CH2:8][CH2:9][N:26]1[C@H:24]([C:21]1[CH:22]=[CH:23][C:18]([Br:17])=[CH:19][CH:20]=1)[CH3:25])=[O:4]Participants
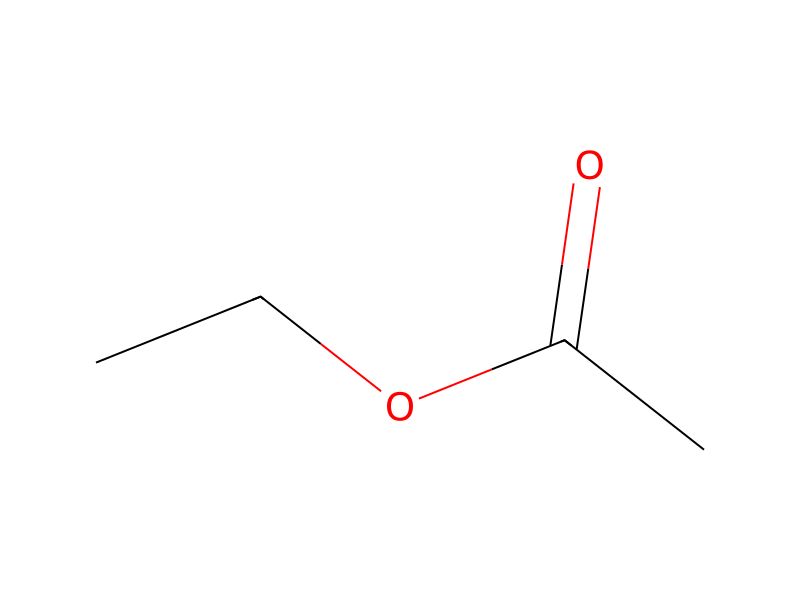
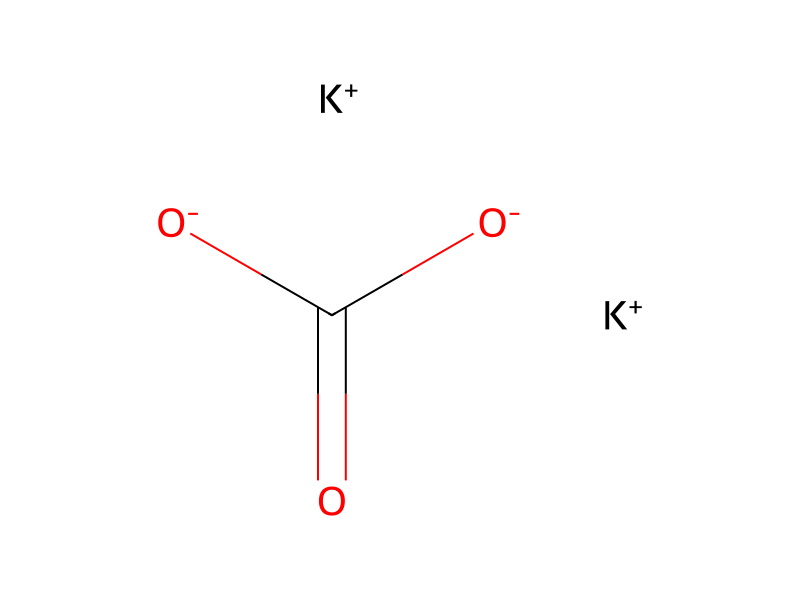



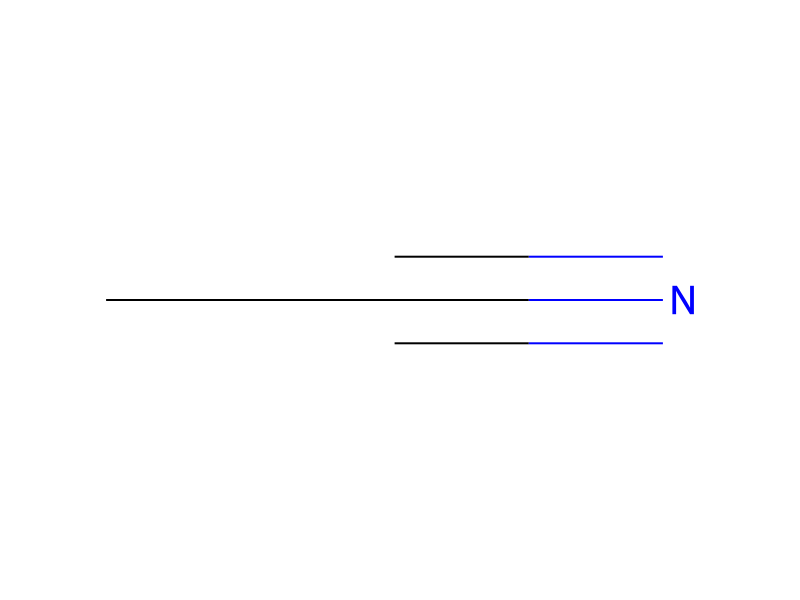
Conditions
- Temperature
- 80 °C
Procedure
The following procedure was adapted from W. A. J. Starmans, R. W. A. Walgers, L. Thijs, R. de Gelder, J. M. M. Smits, and B. Zwanenburg, Tetrahedron 54 (1998) 4991-5004. To a mixture of methyl 2,5-dibromopentanoate (0.401 ml, 2.56 mmol) and K2CO3 (706 mg, 5.11 mmol) in MeCN (7 ml) and water (0.7 ml) was added dropwise a solution of (S)-1-(4-bromophenyl)ethanamine (0.405 ml, 2.81 mmol) in MeCN (3.5 ml) at 80° C. After the addition, the reaction mixture was further stirred at 80° C. for 14 h then cooled to RT, diluted into AcOEt (50 ml) and washed with water (40 ml). The aqueous phase was further extracted with AcOEt and the combined organic fractions were dried over Na2SO4, filtered and evaporated to dryness. The resulting crude material was purified and both diastereoisomers separated by reversed phase prep-HPLC (gradient elution, MeCN/water containing 20 mM ammonium formate). Fractions containing pure material were combined and concentrated under vacuum. The resulting aqueous mixture was basified by the addition of Na2CO3 2M and extracted with AcOEt. The organic layer was washed with brine, dried over Na2SO4, filtered and evaporated to dryness to yield the title compounds as light yellow oils. Intermediate 89.1:240 mg, 0.77 mmol, 30%; HPLC: AtRet=1.15 min, CtRet=4.07 min; LC-MS: m/z 314.4 [M+H]+; 1H NMR (400 MHz, CDCl3): 1.38 (d, J=6.6, 3H), 1.76-1.99 (m, 3H), 2.04-2.16 (m, 1H), 2.52 (q, J=8.1, 1H), 3.05-3.13 (m, 1H), 3.47 (dd, J=9.4, 3.5, 1H), 3.51 (s, 3H), 3.66 (q, J=6.6, 1H), 7.21-7.26 (m, 2H), 7.39-7.45 (m, 2H). Intermediate 89.2: 222 mg, 0.71 mmol, 28%; HPLC: AtRet=1.15 min, CtRet=5.18 min; LC-MS: m/z 314.4 [M+H]+; 1H NMR (400 MHz, CDCl3): 1.36 (d, J=6.8, 3H), 1.73-1.84 (m, 1H), 1.84-1.96 (m, 2H), 2.03-2.14 (m, 1H), 2.54-2.63 (m, 1H), 2.93-3.02 (m, 1H), 3.31 (dd, J=9.2, 4.0, 1H), 3.69 (s, 3H), 3.72 (q, J=6.6, 1H), 7.17-7.22 (m, 2H), 7.42-7.47 (m, 2H). The configuration of each diastereoisomers was attributed by comparing the 1H NMR spectra with published 1H NMR data of close analogues (see R. Almansa, D. Guijarro and M. Yus Tetrahedron: Asymmetry 18, 2007, 2828-2840).
Workup
- additionAfter the addition
- temperaturethen cooled to RT
- washwashed with water (40 ml)
- extractionThe aqueous phase was further extracted with AcOEt
- dry with materialthe combined organic fractions were dried over Na2SO4
- filtrationfiltered
- customevaporated to dryness
- customThe resulting crude material was purified
- customboth diastereoisomers separated by reversed phase prep-HPLC (gradient elution, MeCN/water containing 20 mM ammonium formate)
- additionFractions containing pure material
- concentrationconcentrated under vacuum
- additionThe resulting aqueous mixture was basified by the addition of Na2CO3 2M
- extractionextracted with AcOEt
- washThe organic layer was washed with brine
- dry with materialdried over Na2SO4
- filtrationfiltered
- customevaporated to dryness
Sources & evidence
- ORD record
- ord-01f1e7c5a88b48a4af5ea2405cf7430e
- Source dataset
- uspto-grants-2013_05
- Patent number
- US08440693B2
Reaction notes
Loading…