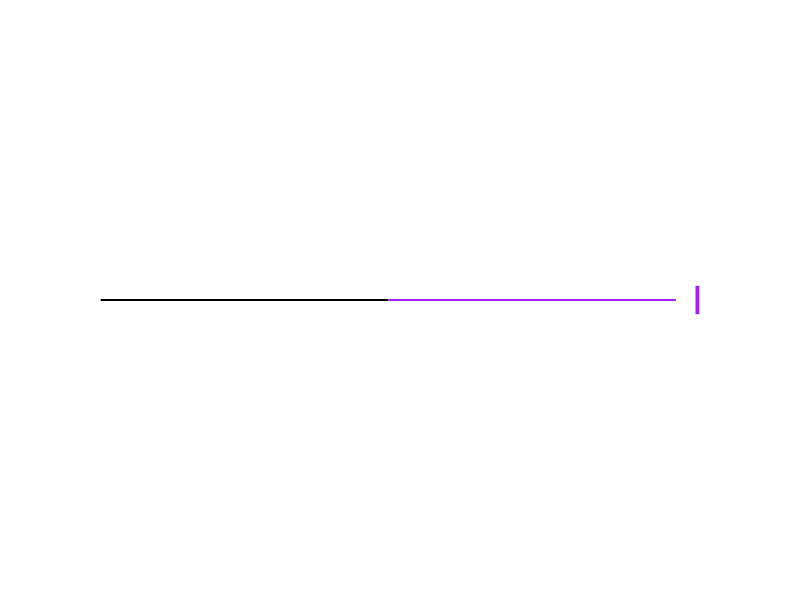
Methyl iodide → 3-Methyl-tetrahydro-pyridazine-1,2,3-tricarboxylic acid 1,2-di-tert-butyl ester 3-methyl ester
Reaction equation

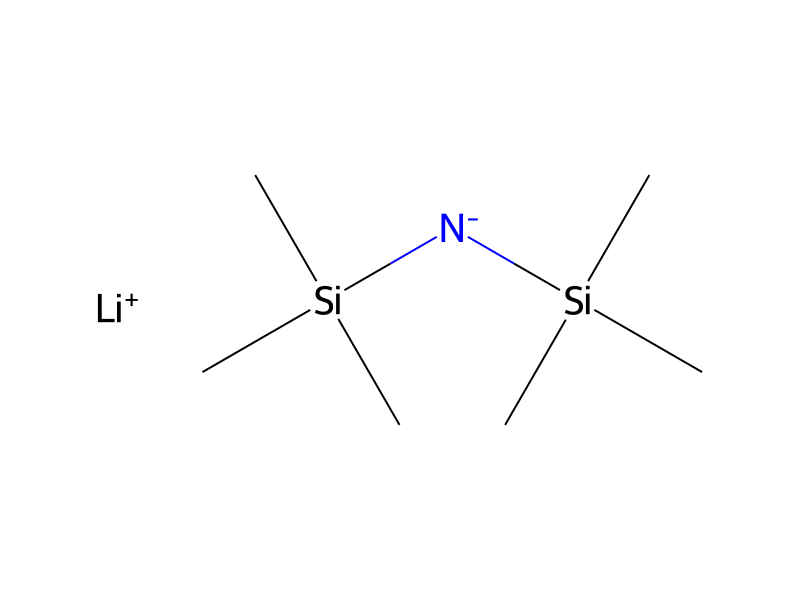
[CH3:1][O:2][C:3]([C@@H:5]1[CH2:10][CH2:9][CH2:8][N:7]([C:11]([O:13][C:14]([CH3:17])([CH3:16])[CH3:15])=[O:12])[N:6]1[C:18]([O:20][C:21]([CH3:24])([CH3:23])[CH3:22])=[O:19])=[O:4].[CH3:25][Si]([N-][Si](C)(C)C)(C)C.[Li+].IC>O1CCCC1>[CH3:1][O:2][C:3]([C:5]1([CH3:25])[CH2:10][CH2:9][CH2:8][N:7]([C:11]([O:13][C:14]([CH3:17])([CH3:15])[CH3:16])=[O:12])[N:6]1[C:18]([O:20][C:21]([CH3:24])([CH3:23])[CH3:22])=[O:19])=[O:4]Participants
Reactant 3

Product 1

Reagents, catalysts & solvents 1

Conditions
- Temperature
- -78 °C
Procedure
To a solution of (S)-tetrahydro-pyridazine-1,2,3-tricarboxylic acid 1,2-di-tert-butyl ester 3-(2,2,2-trichloro-ethyl) ester (5.0 g, 10.8 mmol) in tetrahydrofuran/methanol (50 mL, 1:1) containing 3 Å molecular sieves at 0° C. was added tetra n-butylammonium fluoride (22 mL, 22 mmol, 1 M in tetrahydrofuran). After stirring at room temperature for 24 h, the mixture was filtered through a pad of Celite and the filtrate was evaporated. The residue was partitioned between diethyl ether and saturated sodium hydrogen carbonate solution. The aqueous layer was extracted with diethyl ether. The organic extracts were combined and the solution was filtered through a hydrophobic frit and the solvent was evaporated. The residue was purified by silica gel chromatography eluting with iso-hexanes/ethyl acetate 20:1 to 3:2 to afford (S)-tetrahydro-pyridazine-1,2,3-tricarboxylic acid 1,2-di-tert-butyl ester 3-methyl ester (3.0 g, 82%) as a colorless oil and as a mixture of rotamers. LCMS (m/z) 345.1 [M+H], Tr=2.78 min. A solution of (S)-tetrahydro-pyridazine-1,2,3-tricarboxylic acid 1,2-di-tert-butyl ester 3-methyl ester (3.0 g, 8.72 mmol) in tetrahydrofuran (20 mL) at −78° C. was treated with lithium bis(trimethylsilyl)amide (13 mL, 13.0 mmol, 1 M in tetrahydrofuran). After stirring at −78° C. for 1 h, the mixture was treated with iodomethane (1.4 mL, 22.5 mmol). The reaction mixture was stirred at −78° C. for 1 h, then at 0° C. for 1 h and at room temperature for 1 h. The reaction mixture was re-cooled to 0° C. and quenched with water. The aqueous phase was extracted with dichloromethane (2×) and the organic solution was passed through a hydrophobic frit and the solvent was evaporated. The residue was purified by silica gel chromatography eluting with a gradient of iso-hexanes/ethyl acetate 4:1 to 0:1 to afford the title compound (1.62 g, 52%) as a colorless oil. 1H NMR (300 MHz, CDCl3) δ 1.41-1.52 (m, 18H), 1.53-1.67 (m, 4H), 1.71-1.99 (m, 3H), 3.17-3.51 (m, 1H), 3.70-3.81 (m, 3H), 3.90-4.21 (m, 1H). LCMS (m/z) 359.0 [M+H], Tr=2.90 min.
Workup
- additionas a mixture of rotamers
- customLCMS (m/z) 345.1 [M+H], Tr=2.78 min
- stirringThe reaction mixture was stirred at −78° C. for 1 h
- waitat 0° C. for 1 h
- waitat room temperature for 1 h
- temperatureThe reaction mixture was re-cooled to 0° C.
- customquenched with water
- extractionThe aqueous phase was extracted with dichloromethane (2×)
- customthe solvent was evaporated
- customThe residue was purified by silica gel chromatography
- washeluting with a gradient of iso-hexanes/ethyl acetate 4:1 to 0:1
Sources & evidence
- ORD record
- ord-6483bcbac4804ada940de042b36ae0ea
- Source dataset
- uspto-grants-2015_06
- Patent number
- US09062092B2
Reaction notes
Loading…