1,1,1,3,3,3-Hexafluoro-2-propanol → HCID 89677180, compound
Reaction equation

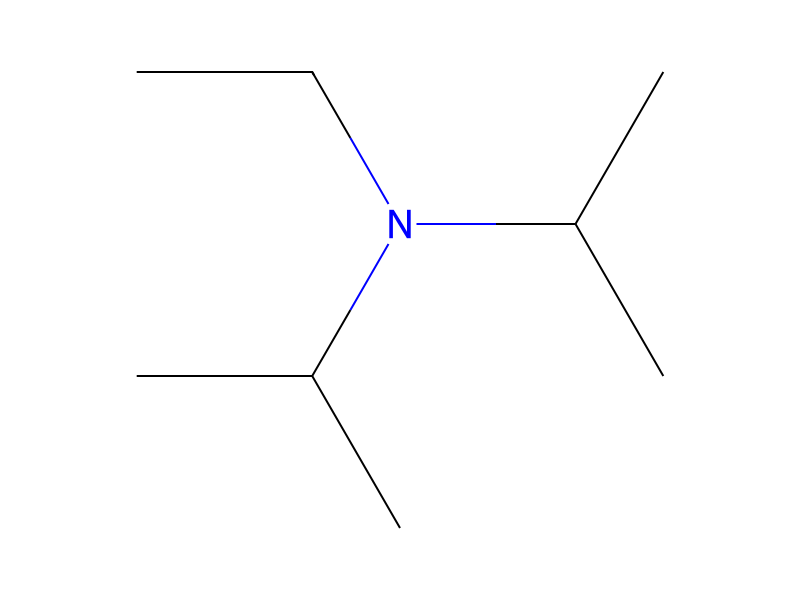
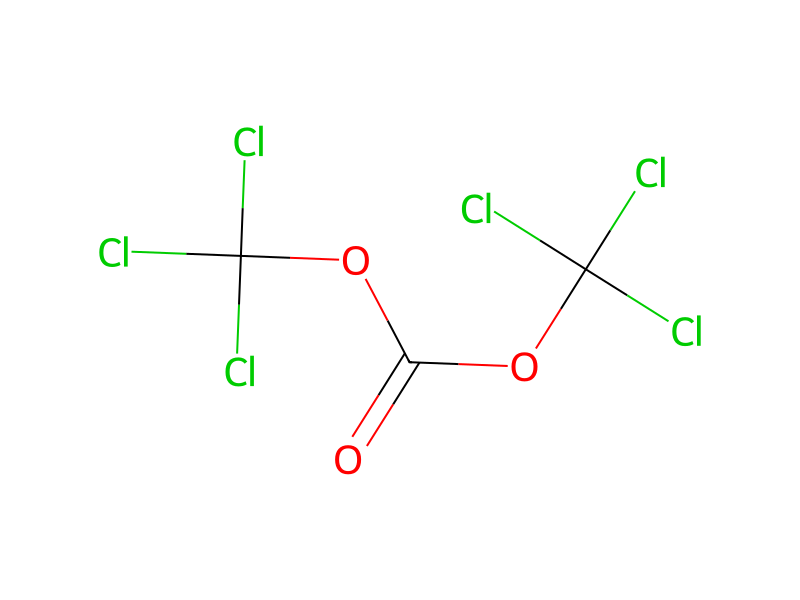
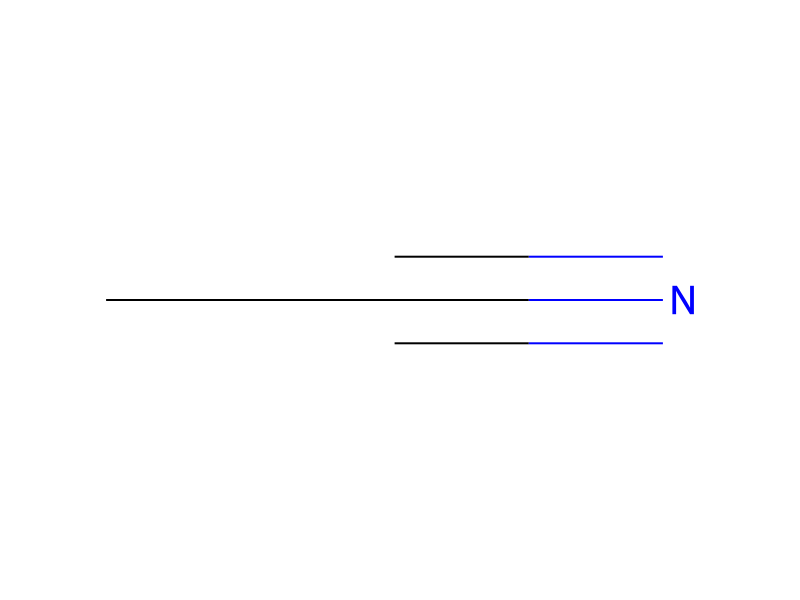
[F:1][C:2]([F:10])([F:9])[CH:3]([OH:8])[C:4]([F:7])([F:6])[F:5].ClC(Cl)(O[C:15](=[O:21])OC(Cl)(Cl)Cl)Cl.C(N(CC)C(C)C)(C)C.[CH3:32][CH:33]1[CH2:38][NH:37][CH2:36][CH2:35][N:34]1[CH2:39][C:40]1[CH:45]=[CH:44][C:43]([C:46]2[CH:51]=[CH:50][CH:49]=[CH:48][CH:47]=2)=[CH:42][CH:41]=1>O.C(#N)C>[CH3:32][CH:33]1[N:34]([CH2:39][C:40]2[CH:45]=[CH:44][C:43]([C:46]3[CH:51]=[CH:50][CH:49]=[CH:48][CH:47]=3)=[CH:42][CH:41]=2)[CH2:35][CH2:36][N:37]([C:15]([O:8][CH:3]([C:4]([F:7])([F:6])[F:5])[C:2]([F:10])([F:9])[F:1])=[O:21])[CH2:38]1Participants




Procedure
A 50 mL round-bottom flask was charged with 1,1,1,3,3,3-hexafluoropropan-2-ol (168 mg, 1.00 mmol, 1.00 equiv), triphosgene (99.0 mg, 0.330 mmol, 0.33 equiv), and acetonitrile (10 mL). N,N-diisopropylethylamine (471 mg, 3.64 mmol, 3.65 equiv) was added dropwise. The mixture was stirred at room temperature for 2 hours. 2-Methyl-1-[(4-phenylphenyl)methyl]piperazine (266 mg, 1.00 mmol, 1.00 equiv) was added. The resulting solution was stirred for 2 hours at room temperature and diluted with water (20 mL). The resulting solution was extracted with dichloromethane (3×20 mL) and the organic layers were dried over anhydrous sodium sulfate, filtered and concentrated under reduced pressure. The residue was chromatographed on a silica gel column with ethyl acetate/petroleum ether (1/3). The product (208 mg) was purified by preparative HPLC using the following gradient conditions: 20% CH3CN/80% Phase A increasing to 80% CH3CN over 10 min, then to 100% CH3CN over 0.1 min, holding at 100% CH3CN for 1.9 min, then reducing to 20% CH3CN over 0.1 min, and holding at 20% for 1.9 min, on a Waters 2767-5 Chromatograph. Column: Xbridge Prep C18, 19*150 mm 5 um; Mobile phase: Phase A: aqueous NH4HCO3 (0.05%); Phase B: CH3CN; Detector, UV220 & 254 nm. Purification resulted in 91.3 mg (20% yield) of 1,1,1,3,3,3-hexafluoropropan-2-yl 3-methyl-4-[(4-phenylphenyl)methyl]piperazine-1-carboxylate as colorless oil. 1H NMR 400 MHz (CDCl3) δ 7.54-7.62 (m, 4H), 7.33-7.49 (m, 5H), 5.75-5.80 (m, 1H), 3.98-4.23 (m, 1H), 3.75-3.86 (m, 2H), 3.01-3.48 (m, 3H), 2.62-2.77 (m, 2H), 2.20-2.27 (m, 1H), 1.19 (br, 3H). LCMS (ESI, m/z): 461 [M+H]+.
Workup
- stirringThe resulting solution was stirred for 2 hours at room temperature
- extractionThe resulting solution was extracted with dichloromethane (3×20 mL)
- dry with materialthe organic layers were dried over anhydrous sodium sulfate
- filtrationfiltered
- concentrationconcentrated under reduced pressure
- customThe residue was chromatographed on a silica gel column with ethyl acetate/petroleum ether (1/3)
- customThe product (208 mg) was purified by preparative HPLC
- wait20% CH3CN/80% Phase A increasing to 80% CH3CN over 10 min
- waitto 100% CH3CN over 0.1 min
- waitholding at 100% CH3CN for 1.9 min
- waitreducing to 20% CH3CN over 0.1 min
- waitholding at 20% for 1.9 min, on a Waters 2767-5 Chromatograph
- customPurification
Sources & evidence
- ORD record
- ord-b799727c148e4ea69ce2f387f664e30d
- Source dataset
- uspto-grants-2015_09
- Patent number
- US09133148B2
Reaction notes
Loading…