2-Furancarbonyl chloride → HCID 67133207, compound
Reaction equation

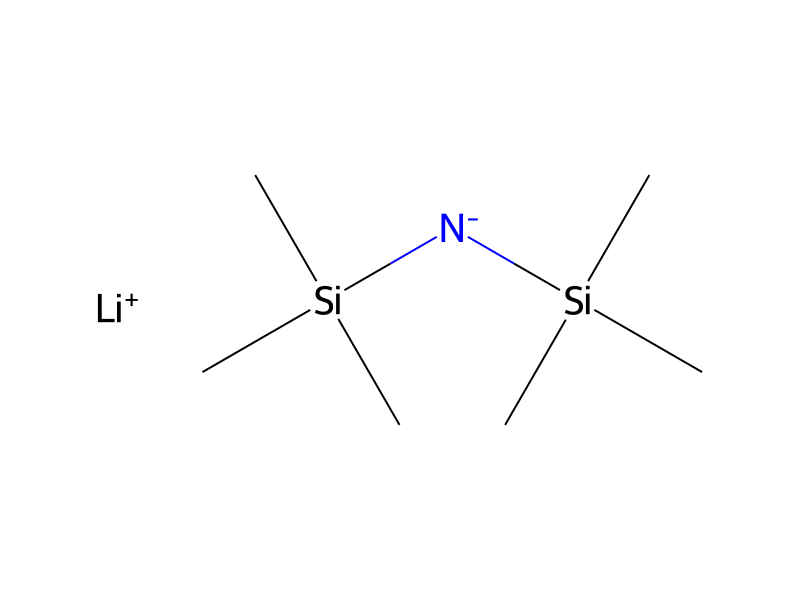
C(OC(=O)C[N:6](C1CC1)C(=O)C1C=CC(OC(F)(F)F)=CC=1)C.[Li+].C[Si]([N-][Si](C)(C)C)(C)C.O1C=CC=C1C(Cl)=O.[CH2:42]([O:44][C:45](=[O:71])[CH:46]([N:54]([CH:68]1[CH2:70][CH2:69]1)[C:55](=O)[C:56]1[CH:61]=[CH:60][C:59]([O:62][C:63]([F:66])([F:65])[F:64])=[CH:58][CH:57]=1)[C:47]([C:49]1[O:50][CH:51]=[CH:52][CH:53]=1)=O)[CH3:43].FC(F)(F)C([O-])=O.[NH4+]>C1COCC1.CCO>[CH2:42]([O:44][C:45]([C:46]1[N:54]([CH:68]2[CH2:70][CH2:69]2)[C:55]([C:56]2[CH:61]=[CH:60][C:59]([O:62][C:63]([F:66])([F:65])[F:64])=[CH:58][CH:57]=2)=[N:6][C:47]=1[C:49]1[O:50][CH:51]=[CH:52][CH:53]=1)=[O:71])[CH3:43]Participants
Reactant 5




Product 1

Reagents, catalysts & solvents 2

Conditions
- Temperature
- -70 °C
Procedure
Under argon, a solution of 1.071 g (3.23 mmol) of [cyclopropyl-(4-trifluoromethoxy-benzoyl)-amino]-acetic acid ethyl ester (intermediate 7B) in 10.0 ml of THF was added drop by drop to 3.5 ml of LiHMDS (solution 1M in THF, 3.5 mmol) cooled to −70° C. The reaction mixture was stirred for 10 min at −70° C. and for 1 hour at −30° C. Then, a solution of 0.167 g (1.28 mmol) of furan-2-carbonyl chloride in 10.0 ml of THF was added drop wise at −30° C. and the reaction mixture was stirred for 1 hour at −30° C. and then allowed to reach RT over 2 hours. The brown solution was partitioned between a sat. NH4Cl solution and EtOAc. The organic phases were washed with brine and the aqueous phases were extracted with EtOAc. The organic layers were dried over magnesium sulfate, filtered and concentrated in vacuo. The residue was purified by column chromatography (EtOAc/n-heptane 1:1) to give 0.3 g of a 1:2 mixture of [cyclopropyl-(4-trifluoromethoxy-benzoyl)-amino]-acetic acid ethyl ester and 2-[cyclopropyl-(4-trifluoromethoxy-benzoyl)-amino]-3-furan-2-yl-3-oxo-propionic acid ethyl ester. This mixture (0.30 g, max. 0.47 mmol) and 0.368 g (2.81 mmol) of ammoniumtrifluoroacetate were stirred for 10 min in 5 ml EtOH at room temperature; then, the solvent was evaporated and the brown oil was heated at 110° C. over night. The reaction mixture was cooled to RT and partitioned between CH2Cl2 and sat. NaHCO3 solution, the aqueous phase was extracted with CH2Cl2. The organic layers were dried over magnesium sulfate, evaporated and chromatographed (silica gel, EtOAc/n-heptane 1:4) to afford 0.098 g (49%) of the title compound as white powder. MS: 407.2 (MH+).
Workup
- stirringthe reaction mixture was stirred for 1 hour at −30° C.
- waitto reach RT over 2 hours
- customThe brown solution was partitioned between a sat. NH4Cl solution and EtOAc
- washThe organic phases were washed with brine
- extractionthe aqueous phases were extracted with EtOAc
- dry with materialThe organic layers were dried over magnesium sulfate
- filtrationfiltered
- concentrationconcentrated in vacuo
- customThe residue was purified by column chromatography (EtOAc/n-heptane 1:1)
- customto give
- customthen, the solvent was evaporated
- temperaturethe brown oil was heated at 110° C. over night
- temperatureThe reaction mixture was cooled to RT
- custompartitioned between CH2Cl2 and sat. NaHCO3 solution
- extractionthe aqueous phase was extracted with CH2Cl2
- dry with materialThe organic layers were dried over magnesium sulfate
- customevaporated
- customchromatographed (silica gel, EtOAc/n-heptane 1:4)
Sources & evidence
- ORD record
- ord-32ea6206ae274e5f8de74b74888c4fbf
- Source dataset
- uspto-grants-2011_05
- Patent number
- US07935707B2
Reaction notes
Loading…