Methyl iodide → Methyl 4-bromo-2-nitrobenzoate
Reaction equation

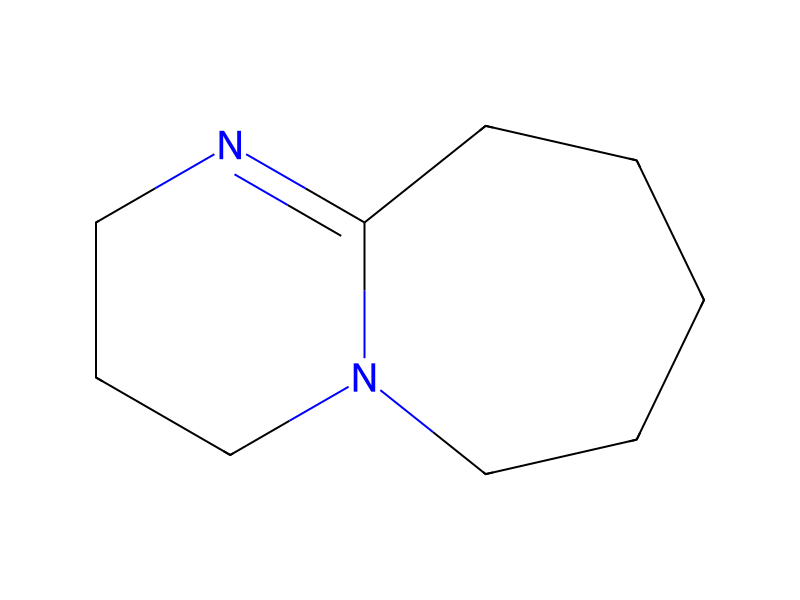
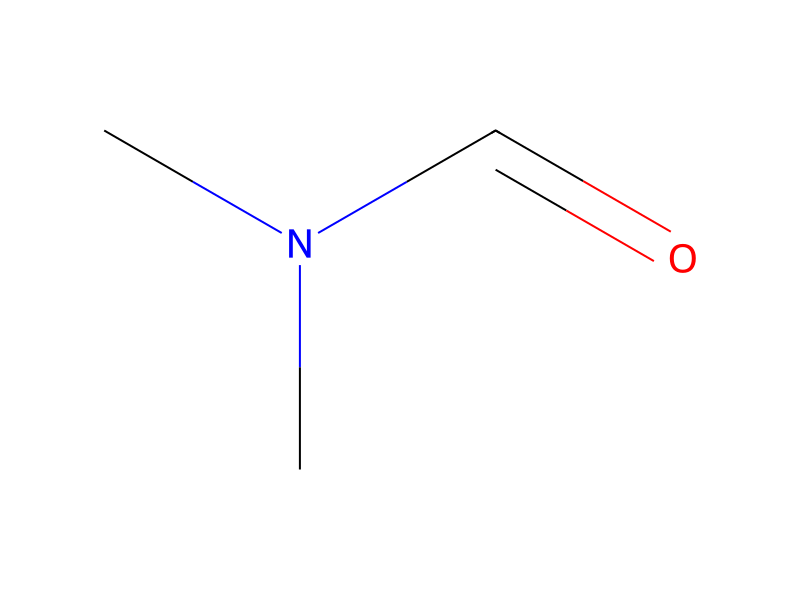
N[C:2]1C=C(Br)C=CC=1C(OC)=O.[Br:13][C:14]1[CH:22]=[CH:21][C:17]([C:18]([OH:20])=[O:19])=[C:16]([N+:23]([O-:25])=[O:24])[CH:15]=1.N12CCCN=C1CCCCC2.IC>CN(C=O)C.O>[Br:13][C:14]1[CH:22]=[CH:21][C:17]([C:18]([O:20][CH3:2])=[O:19])=[C:16]([N+:23]([O-:25])=[O:24])[CH:15]=1Participants
Reactant 4


Product 1

Reagents, catalysts & solvents 2

Conditions
- Temperature
- 0 °C
Procedure
Diethylthiocarbamic acid O-quinoxalin-5-yl ester. A mixture of 5-hydroxyquinoxaline (2.13 g, 14.6 mmol), finely ground K2CO3 (4.0 g, 29 mmol), and DMF (50 mL) was stirred at 23° C. for 1 h. Solid diethylthiocarbamoyl chloride (2.43 g, 16.1 mmol) was then added in a single portion. The resulting mixture was stirred for 2 h, then was diluted with H2O (150 mL) and extracted with diethyl ether (2×100 mL). The combined ethereal extracts were washed with H2O (100 mL) and brine (100 mL), then dried and concentrated to a viscous orange oil, which was used without purification in the subsequent step (3.63 g, 95%). MS (ESI): Calculated for C13H15N3OS, 261.1; found, m/z 262 [M+H]+. 1H NMR (500 MHz, CDCl3): 8.85-8.65 (m, 2H), 7.96 (dd, J=8.5, 1.1 Hz, 1H), 7.71 (t, J=7.9 Hz, 1H), 7.46 (dd, J=7.6, 1.18 Hz, 1H), 3.87 (q, J=7.1 Hz, 2H), 3.78 (q, J=7.1 Hz, 2H), 1.38 (t, J=7.1 Hz, 3H), 1.28 (t, J=7.1 Hz, 3H). 13C NMR (125 MHz, CDCl3): 186.6, 149.4, 144.9, 144.5, 143.4, 137.0, 128.9, 127.0, 123.1, 48.2, 44.5, 13.1, 11.5. B. Diethylthiocarbamic acid S-guinoxalin-5-yl ester. Neat diethylthiocarbamic acid O-quinoxalin-5-yl ester (0.52 g, 2.0 mmol) was heated to 240° C. for 1 h. The resulting brown oil was chromatographed (20 to 50% EtOAc/hexanes), providing a pale yellow oil (0.49 g, 94%). MS (ESI): Calculated for C13H15N3OS, 261.1; found, m/z 262 [M+H]+. 1H NMR (500 MHz, CDCl3): 8.93 (d, J=1.8 Hz, 1H), 8.87 (d, J=1.8 Hz, 1H), 8.18 (dd, J=8.4, 1.2 Hz, 1H), 8.13 (dd, J=7.3, 1.2 Hz, 1H), 7.81 (dd, J=7.3, 1.0 Hz, 1H), 3.61 (br s, 2H), 3.43 (br s, 2H), 1.38 (br s, 3H), 1.16 (br s, 3H). C. Quinoxaline-5-sulfonyl chloride. A solution of diethylthiocarbamic acid S-quinoxalin-5-yl ester (3.20 g, 12.3 mmol), KOH (6.89 g, 123 mmol) and methanol (100 mL) was heated at reflux for 16 h. The solution was allowed to cool to 23° C., and then AcOH (7 mL) was added. The mixture was diluted with H2O (100 mL) and extracted with EtOAc (2×100 mL). The combined extracts were washed with H2O (100 mL) and brine (100 mL), then dried and concentrated to a tan solid (1.90 g). A portion of this thiol (0.22 g, 1.4 mmol) was combined with DCM (50 mL), formic acid (25 mL), and H2O (25 mL), and the resulting biphasic mixture was cooled to 0° C. Chlorine gas was bubbled through this mixture with rapid stirring for 5 min. The mixture was transferred to a separatory funnel, and the organic phase was collected. The aqueous phase was extracted with DCM (50 mL), and the combined organic phases were washed with 1 M NaOH (50 mL) and brine (50 mL), then dried. The solution was concentrated to afford the titled compound as a light yellow crystalline solid (0.28 g, 86%). 1H NMR (500 MHz, CDCl3): 9.17 (d, J=1.8 Hz, 1H), 9.07 (d, J=1.8 Hz, 1H), 8.60 (dd, J=7.5, 1.4 Hz, 1H), 8.53 (dd, J=8.4, 1.4 Hz, 1H), 7.96 (dd, J=8.6, 0.8 Hz, 1H). 13C NMR (125 MHz, CDCl3): 146.9, 146.8, 143.7, 140.4, 139.0, 138.4, 132.4, 128.8. D. 4-Bromo-2-nitrobenzoic acid. A mixture of 4-bromo-2-nitrotoluene (5.0 g, 23 mmol), KMnO4 (1 g, 70 mmol), and H2O (250 mL) was heated at reflux overnight in a 1 L round-bottom flask fitted with a reflux condenser. The brown suspended MnO2 was removed by filtration through a pad of diatomaceous earth. The filter cake was washed with H2O. The basic filtrate was acidified to pH ˜1 with concentrated HCl and extracted with EtOAc (3×300 mL). The combined organic layers were dried (MgSO4) and concentrated in vacuo to provide the pure benzoic acid (1.22 g, 22%). MS (ESI) calculated for C7H4BrNO4, 244.9; found, m/z 244 [M−H]−. 1H NMR (400 MHz, CD3OD): 8.07 (d, J=1.9 Hz, 1H), 7.85 (dd, J=8.2, 1.9 Hz, 1H), 7.65 (d, J=8.2 Hz, 1H). E. Methyl 2-amino-4-bromobenzoate. To a stirred solution of 4-bromo-2-nitrobenzoic acid (3.8 g, 15 mmol) in DMF (30 mL) at 0° C. was added 1,8-diazabicyclo[5.4.0]undec-7-ene (DBU) (10.0 mL, 75.0 mmol) followed by iodomethane (4.7 mL, 75 mmol). The reaction mixture was stirred 15 min at 0° C., then was allowed to warm to room temperature and was stirred overnight. The mixture was poured into H2O and extracted with EtOAc (2×). The combined organic extracts were washed with H2O (2×), dried (MgSO4), and concentrated in vacuo. The residue was purified by flash chromatography (hexanes/EtOAc) to afford methyl 4-bromo-2-nitrobenzoate as a pale yellow solid (3.52 g, 0.90%). To a solution of the nitrobenzoate (3.52 g, 13.5 mmol) in 1:1 EtOAc/DCM (30 mL) at room temperature was added SnCl2.2H2O (15 g, 67 mmol). The reaction mixture was allowed to stir overnight. The solvents were evaporated in vacuo, and the residue was partitioned between satd. aq. NaHCO3 and DCM. The layers were separated, and the aqueous layer was further extracted with DCM (2×). The combined organic layers were dried (MgSO4) and concentrated in vacuo to provide the pure aminobenzoate as a white solid (2.89 g, 93%). 1H NMR (400 MHz, CDCl3): 7.70 (d, J=8.6 Hz, 1H), 6.84 (d, J=1.9 Hz, 1H), 6.75 (dd, J=8.6, 1.9 Hz, 1H), 5.78 (br s, 2H), 3.86 (s, 3H). F. 4-Bromo-2-(quinoxaline-5-sulfonylamino)benzoic acid methyl ester. A solution of quinoxaline-5-sulfonyl chloride (0.50 g, 2.2 mmol), methyl 2-amino-4-bromobenzoate (0.50 g, 2.2 mmol), pyridine (0.87 mL, 11 mmol), and DCM (15 mL) was maintained at 23° C. for 16 h. The reaction mixture was diluted with EtOAc (50 mL) and washed with satd. aq. NaHCO3 (50 mL), then dried and concentrated. The residue was purified by flash chromatography (5 to 40% EtOAc/hexanes) to afford the sulfonamide as a white solid (0.78 g, 84%). MS (ESI) calculated for C16H12BrN3O4S, 421.0; found, m/z 422 [M+H]+. 1H NMR (500 MHz, CDCl3): 11.39 (s, 1H), 8.96 (d, J=1.7 Hz, 1H), 8.94 (d, J=1.7 Hz, 1H), 8.61 (dd, J=7.4, 1.4 Hz, 1H), 8.33 (dd J=8.5, 1.4 Hz, 1H), 8.02 (d, J=1.9 Hz, 1H), 7.89 (dd, J=8.4, 1.0 Hz, 1H), 7.69 (d, J=8.5, 1H), 7.06 (dd, J=8.5, 1.9 Hz, 1H), 3.90 (s, 3H). G. 4-Bromo-2-(quinoxaline-5-sulfonylamino)benzoic acid. A solution of LiOH.H2O (0.36 g, 8.6 mmol) in H2O (5 mL) was added to a solution of 4-bromo-2-(quinoxaline-5-sulfonylamino)benzoic acid methyl ester (0.73 g, 1.7 mmol) and THF (10 mL), and the biphasic mixture was rapidly stirred for 16 h. The mixture was concentrated to a volume of 5 mL, and then was adjusted to pH 5 with 1 M HCl. The resulting precipitate was collected by filtration, providing the acid as a white solid (0.68 g, 96%). MS (ESI) calculated for C15H10BrN3O4S, 407.0; found, m/z 408 [M+H]+. 1H NMR (500 MHz, CDCl3): 14.1 (br s, 1H), 9.11 (d, J=1.4 Hz, 1H), 8.99 (d, J=1.4 Hz, 1H), 8.63 (d, J=7.4 Hz, 1H), 8.42 (d, J=8.4 Hz, 1H), 8.06 (t, J=8.3 Hz, 1H), 7.77 (d, J=1.6 Hz, 1H), 7.71 (d, J=8.5 Hz, 1H), 7.20 (dd, J=8.5, 1,6 Hz, 1H). H. S-(S)-2-Methyl-propane-2-sulfinic acid 2,4-dichloro-benzylideneamide. A suspension of 2,4-dichlorobenzaldehyde (0.75 g, 4.3 mmol), (S)-tert-butanesulfinamide (0.47 g, 3.9 mmol), and powdered anhydrous CuSO4 (1.2 g, 7.8 mmol) in DCM (8 mL) was stirred overnight. The reaction mixture was filtered, and the filter cake was washed with DCM. The filtrate was concentrated in vacuo to give the crude N-sulfinyl imine as white solid. Purification by flash chromatography (EtOAc/hexanes) provided 0.97 g (90%) of the N-sulfinyl imine as a white solid. 1H NMR (400 MHz, CDCl3): 8.98 (s, 1H), 8.01 (d, J=8.5 Hz, 1H), 7.48 (d, J=2.1 Hz, 1H), 7.35-7.32 (m, 1H), 1.27 (s, 9H). I. S-(S)-2-Methyl-propane-2-sulfinic acid 1-(R)-[1-(2,4-dichloro-Phenyl)-ethyl]-amide. To a stirred solution of the above N-sulfinyl imine (0.97 g, 3.5 mmol) in DCM (20 mL) at −50° C. was added a solution of methyl magnesium bromide (3.0 M in diethyl ether, 2.3 mL, 6.9 mmol). The reaction mixture was stirred at −50° C. for 1 h then allowed to warm slowly to room temperature overnight. The reaction was quenched by the addition of satd. aq. NH4Cl, and the mixture was poured into H2O and extracted with DCM (3×). The combined organic layers were dried (Na2SO4) and concentrated in vacuo. Purification by flash chromatography (EtOAc/hexanes) provided the title compound as a colorless solid (1.02 g, 99%, 76% de). Major diastereomer: 1H NMR (400 MHz, CDCl3): 7.43-7.35 (m, 2H), 7.26-7.23 (m, 1H), 5.01 (dq, J=6.7, 4.0 Hz, 1H), 3.39 (d, J=3.7 Hz, 1H), 1.53 (d, J=6.7 Hz, 3H), 1.21 (s, 9H). J. (R)-1-(2,4-Dichlorophenyl)-ethylamine hydrochloride. To a stirred solution of the above sulfinamide (76% de, 1.02 g, 3.47 mmol) in 7:4 methanol/DCM (11 mL) at room temperature was added 2 mL of a satd. solution of HCl (g) in methanol. After several minutes, precipitated amine hydrochloride was visible. The reaction mixture was allowed to stir for 2 h at room temperature. The heterogeneous mixture was concentrated in vacuo until approximately 2 mL remained, and then the amine hydrochloride was fully precipitated by the addition of diethyl ether (10 mL). The HCl salt was collected by suction filtration, washed with diethyl ether, and dried in vacuo to give fine white crystals (722 mg, 92%, 76% ee). 1H NMR (400 MHz, CD3OD): 7.62 (d, J=2.2 Hz, 1H), 7.57 (d, J=8.4 Hz, 1H), 7.50 (dd, J=8.4, 2.2 Hz, 1H), 1.62 (d, J=6.8 Hz, 3H). K. (R)-4-Bromo-N-[1-(2,4-dichlorophenyl)-ethyl]-2-(quinoxaline-5-sulfonylamino)-benzamide. To a solution of 4-bromo-2-(quinoxaline-5-sulfonylamino)benzoic acid (0.021 g, 0.051 mmol) in a mixture of THF (0.08 mL) and DMF (0.40 mL) at room temperature was added pyridine (0.012 mL, 0.15 mmol) followed by O-(7-azabenzotriazol-1-yl)-N,N,N′,N′-tetramethyluronium hexafluorophosphate (HATU) (0.038 g, 0.12 mmol). The reaction mixture was agitated for 1 h on a shaker. (R)-1-(2,4-Dichlorophenyl)-ethylamine hydrochloride (0.038 g, 0.10 mmol) and N,N-diisopropylethylamine (Hünig's base) (0.017 mL, 0.10 mmol) were added. The reaction mixture was agitated for 1 h. TFA (0.050 mL) was added to quench the reaction. The mixture was diluted with DMF (1 mL), and the product amide was obtained by purification of the resulting mixture by preparative reverse-phase chromatography. The title amide was obtained as a solid (24 mg, 83%). MS (ESI): mass calculated for C23H17BrCl2N4O3S, 577.96; m/z found, 577/579/581 [M−H]−. HPLC (reverse phase): RT═10.34 min. 1H NMR (500 MHz, CDCl3): 11.32 (s, 1H), 8.85(d, J=1.8 Hz, 1H), 8.76 (d, J=1.8 Hz, 1H), 8.55 (dd, J=7.4, 1.4 Hz, 1H), 8.31 (dd, J=7.4, 1.4 Hz, 1H), 7.94 (d, J=1.8 Hz, 1H), 7.87 (dd, J=8.4, 7.4 Hz, 1H), 7.44 (d, J=1.7 Hz, 1H), 7.35-7.15 (m, 3H), 7.09 (dd, J=8.4, 1.8 Hz, 1H), 6.36 (br d, J=6.5 Hz, 1H), 5.49-5.30 (m, 1H), 1.52 (d, J=7.0 Hz, 3H).
Workup
- temperatureto warm to room temperature
- stirringwas stirred overnight
- extractionextracted with EtOAc (2×)
- washThe combined organic extracts were washed with H2O (2×)
- dry with materialdried (MgSO4)
- concentrationconcentrated in vacuo
- customThe residue was purified by flash chromatography (hexanes/EtOAc)
Sources & evidence
- ORD record
- ord-f5f4b693d34a4afc9a66ff2ae07ae78d
- Source dataset
- uspto-grants-2007_12
- Patent number
- US07304051B2
Reaction notes
Loading…