Reaction equation

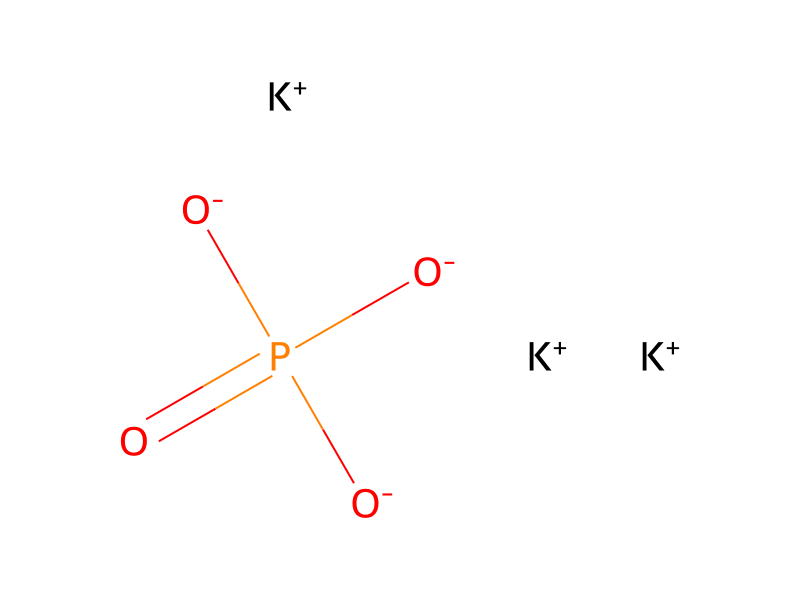
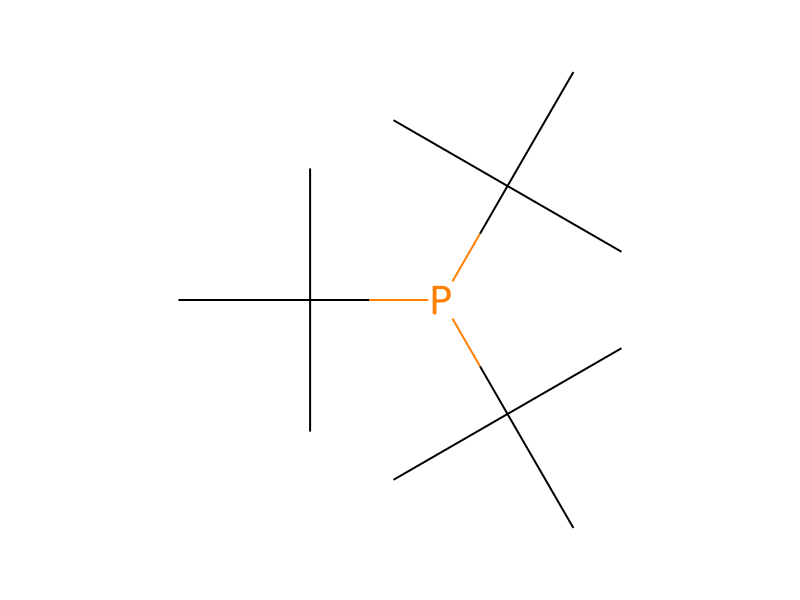
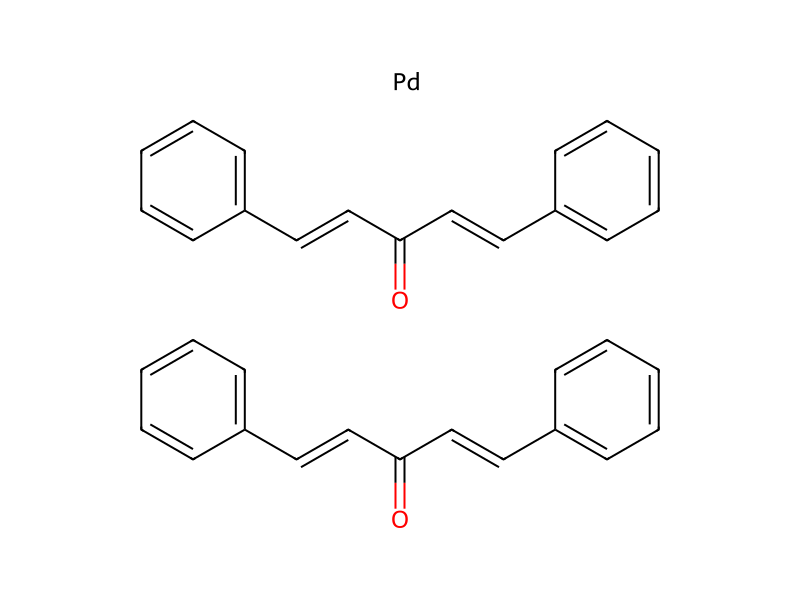
[F:1][CH:2]([F:10])[C:3]1[CH:4]=[C:5](Br)[CH:6]=[CH:7][CH:8]=1.C1(C(C2C=CC=CC=2)=[N:18][CH2:19][C:20]([O:22][CH2:23][CH3:24])=[O:21])C=CC=CC=1.C(P(C(C)(C)C)C(C)(C)C)(C)(C)C.P([O-])([O-])([O-])=O.[K+].[K+].[K+].Cl>C1(C)C=CC=CC=1.C1C=CC(/C=C/C(/C=C/C2C=CC=CC=2)=O)=CC=1.C1C=CC(/C=C/C(/C=C/C2C=CC=CC=2)=O)=CC=1.[Pd]>[NH2:18][CH:19]([C:5]1[CH:6]=[CH:7][CH:8]=[C:3]([CH:2]([F:10])[F:1])[CH:4]=1)[C:20]([O:22][CH2:23][CH3:24])=[O:21]Participants
Reactant 5


Product 1

Reagents, catalysts & solvents 2
Procedure
Quantities of 1.0 g (4.83 mmol) of 3-(difluoromethyl)phenyl bromide, 1.42 g (5.31 mmol) of ethyl N-(diphenylmethylene)glycinate, 0.19 ml (0.193 mmol) of a 1M solution of tri-tert-butylphosphane in toluene, 55 mg (0.10 mmol) of bis(dibenzylideneacetone)palladium(0), 3.08 g (14.49 mmol) of potassium phosphate and 6.04 ml (18.11 mmol) of 3M hydrochloric acid were heated under argon to 100° C. in 20 ml of degassed toluene, and stirred at this temperature overnight. A further 0.19 ml (0.193 mmol) of tri-tert-butylphosphane (1M solution in toluene) and 55 mg (0.10 mmol) of bis(dibenzylideneacetone)palladium(0) were added and the mixture was stirred at 100° C. for a further 24 h. The mixture was cooled to RT and filtered over celite. The celite was washed with a little toluene and the filtrate was freed from the solvent under reduced pressure. For elimination of the protecting group, the residue was taken up in 50 ml of acetonitrile and admixed with 15 ml of 3M hydrochloric acid. After 2 h, the acetonitrile fraction was removed on a rotary evaporator. The aqueous residue was diluted with water to a volume of approximately 150 ml and washed three times with diethyl ether. The aqueous phase was adjusted to a pH of 9 using 2M aqueous sodium carbonate solution and then extracted three times with dichloromethane. The combined organic phases were dried over sodium sulphate, filtered and freed from the solvent on a rotary evaporator. Drying in an HV gave 265 mg (6.46% of theory) of the title compound, in a purity of about 27%, which was reacted further without additional purification.
Workup
- stirringthe mixture was stirred at 100° C. for a further 24 h
- filtrationfiltered over celite
- washThe celite was washed with a little toluene
- waitAfter 2 h
- customthe acetonitrile fraction was removed on a rotary evaporator
- additionThe aqueous residue was diluted with water to a volume of approximately 150 ml
- washwashed three times with diethyl ether
- extractionextracted three times with dichloromethane
- dry with materialThe combined organic phases were dried over sodium sulphate
- filtrationfiltered
- customfreed from the solvent on a rotary evaporator
- customDrying in an HV
Sources & evidence
- ORD record
- ord-de269cd8a1e64422b81b17c84a48f0f1
- Source dataset
- uspto-grants-2014_08
- Patent number
- US08796324B2
Linked notes
Loading…