Reaction equation

B.O1[CH2:6][CH2:5][CH2:4]C1.[N:7]1[CH:12]=[CH:11]C=[CH:9][CH:8]=1.B.B.[CH3:15]SC.S(C)C.[B]>N1C=CC=CC=1.C1COCC1>[CH3:11][CH2:12][N:7]([CH:8]([CH3:9])[CH3:15])[CH:5]([CH3:6])[CH3:4]Participants


Procedure
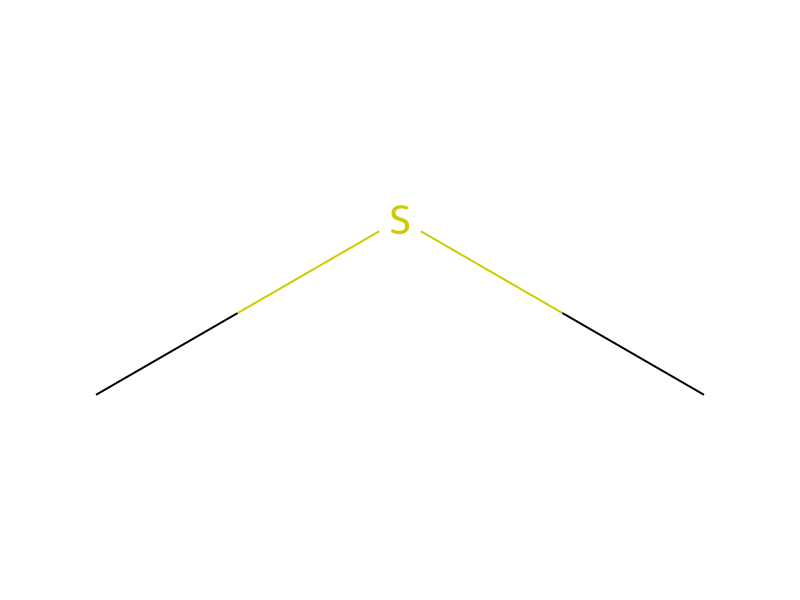
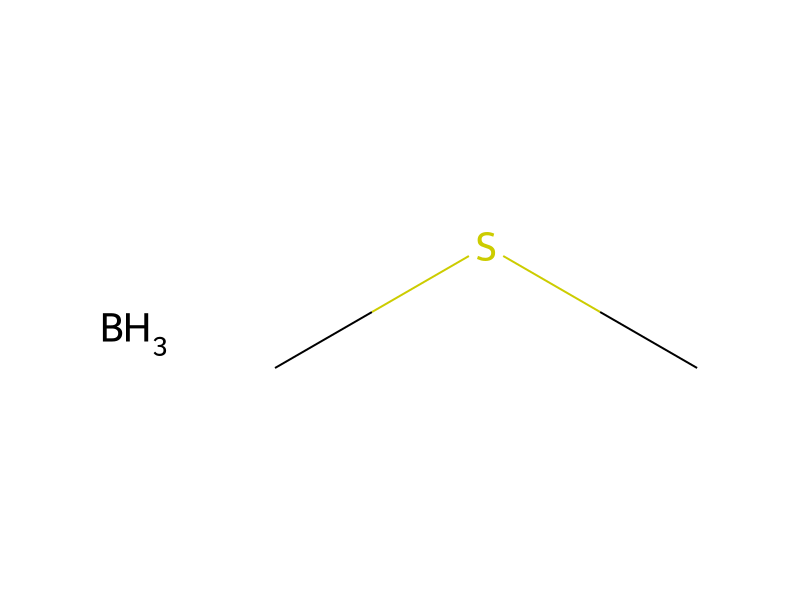
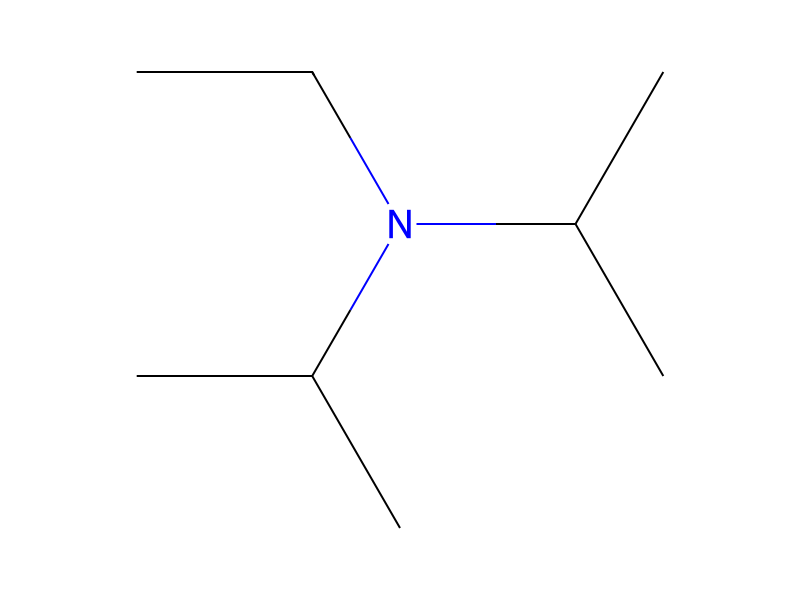
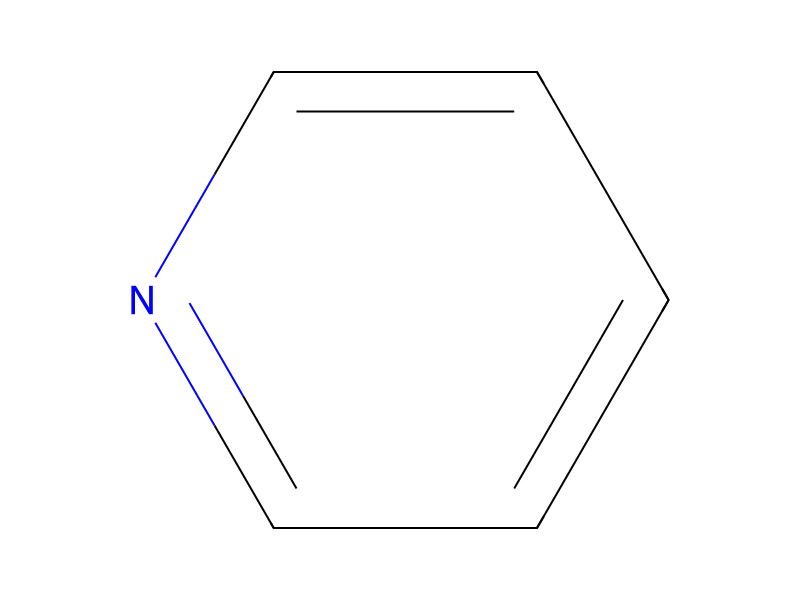
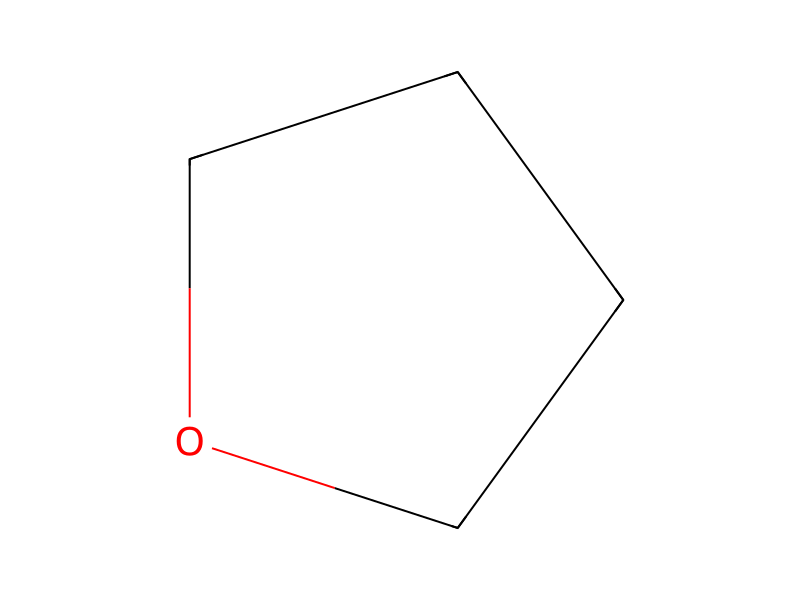
As the phosphorous atom of the H-phosphinate intermediate does not present a pair of free electrons and is not a good donor for the introduction of the BH3 group, the boronation procedure requires an in situ intermediate activation of the H-phosphinate with a silylation agent to obtain a disilyl-phosphonite. The H-phosphinates 5 and 10 are activated in situ in anhydrous THF with N,O-bis(trimethylsilyl)acetamide (BSA) for 1 hr. in their corresponding disilyl-phosphonite intermediates. Various boron complexes have been tested in different solvents to optimise the boronation conditions. A rapid and effective boronation was finally obtained with the complex borane-N,N-diisopropylethylamine (BH3:DIPEA). The complexes borane-tetrahydrofuran (BH3:THF), borane-pyridine (BH3:Pyridine), and borane-dimethylsulfide (BH3:Me2S) require significant reduction times and do not improve the yield. In addition, in order to reduce the formation of diboranated products, the quantity of boron complexes was reduced to a maximum of 2 equivalents to obtain the best compromise between consumption of initial material and the appearance of by-products. In situ boronation of the disilyl phosphonites of 5 and 10 with 2 equivalents of BH3:DIPEA results in the formation of -boranophosphonate intermediates. Without isolation, the intermediates obtained were treated with 30% concentrated ammonium hydroxide in methanol (1:1, v/v) to cleave the trimethylsilyl groups thus making it possible to obtain α-boranophosphonates 6a and 6b. The presence of the P—B bond was confirmed by 31P NMR showing a typical peak at 83 ppm and 98 ppm respectively. α-boranophosphonates 6a and 6b are purified by reverse-phase chromatography with a yield of 28% and 32% respectively.
Workup
- customto obtain the best compromise
- custombetween consumption of initial material
Sources & evidence
- ORD record
- ord-38a6062b2b724ceca260890ebb78b42e
- Source dataset
- uspto-grants-2014_11
- Patent number
- US08884011B2
Linked notes
Loading…