Acetic Acid → HCID 73774836, compound
Reaction equation

[CH:1]([CH:3]1[CH2:8][CH2:7][CH2:6][CH2:5][N:4]1C(OC(C)(C)C)=O)=O.[NH2:16][C:17]1[CH:22]=[CH:21][C:20]([C:23]2[CH:27]=[C:26]([C:28]3[C:29]([N:46](C(OC(C)(C)C)=O)C(=O)OC(C)(C)C)=[N:30][CH:31]=[C:32]([C:34]4[CH:39]=[CH:38][C:37]([S:40]([CH:43]([CH3:45])[CH3:44])(=[O:42])=[O:41])=[CH:36][CH:35]=4)[N:33]=3)[O:25][N:24]=2)=[CH:19][CH:18]=1.C(O)(=O)C.C(O[BH-](OC(=O)C)OC(=O)C)(=O)C.[Na+].C(O)(C(F)(F)F)=O>C(Cl)Cl.C(=O)([O-])O.[Na+]>[CH:43]([S:40]([C:37]1[CH:36]=[CH:35][C:34]([C:32]2[N:33]=[C:28]([C:26]3[O:25][N:24]=[C:23]([C:20]4[CH:19]=[CH:18][C:17]([NH:16][CH2:1][CH:3]5[CH2:8][CH2:7][CH2:6][CH2:5][NH:4]5)=[CH:22][CH:21]=4)[CH:27]=3)[C:29]([NH2:46])=[N:30][CH:31]=2)=[CH:39][CH:38]=1)(=[O:41])=[O:42])([CH3:45])[CH3:44]Participants
Reactant 5
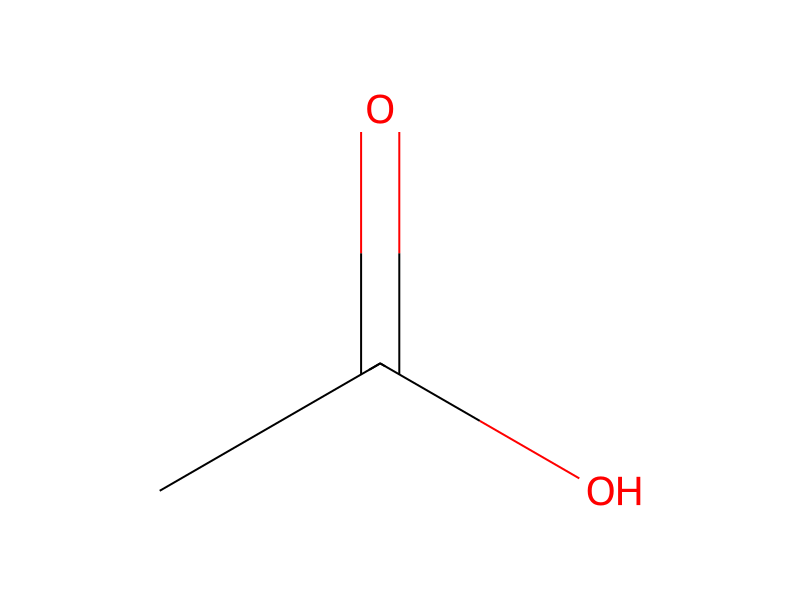

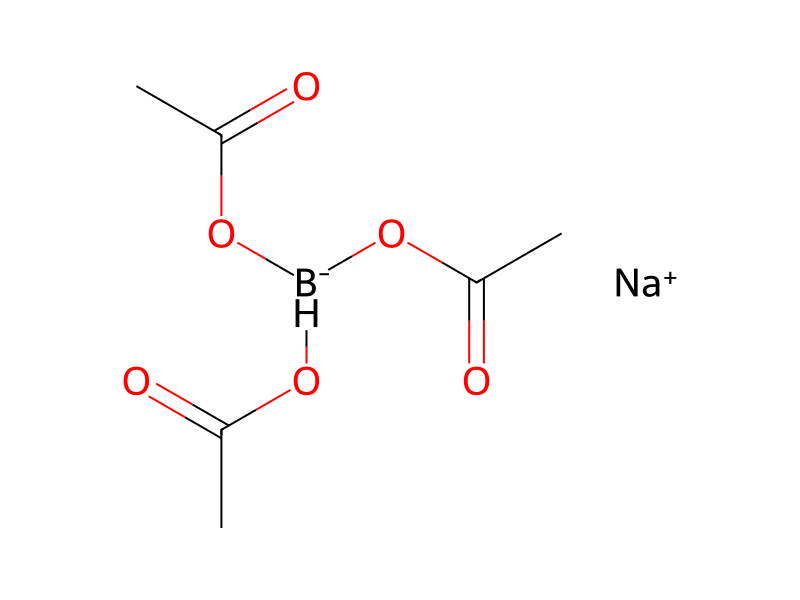


Product 1

Reagents, catalysts & solvents 2

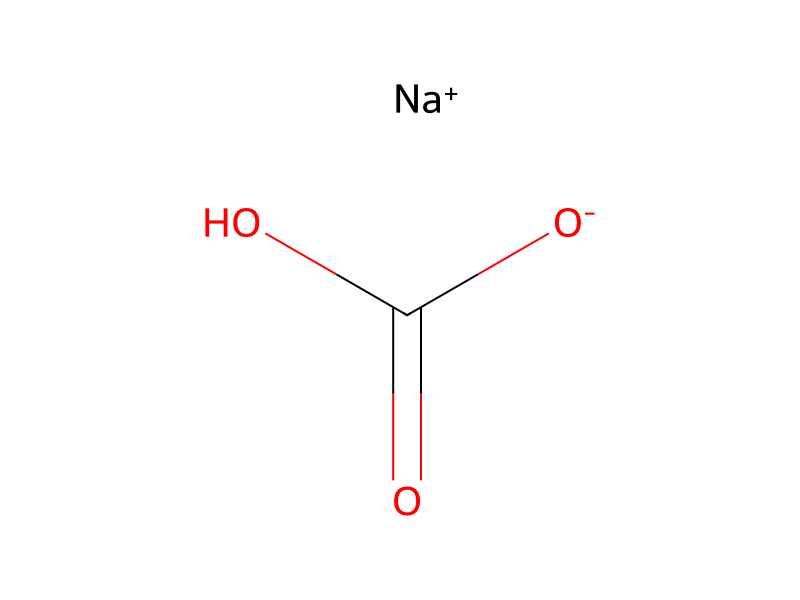
Conditions
- Temperature
- 5 °C
Procedure
To a solution of tert-butyl 2-formylpiperidine-l-carboxylate (18.45 mg, 0.08652 mmol) and tert-butyl N-[3-[3-(4-aminophenyl)isoxazol-5-yl]-5-(4-isopropylsulfonylphenyl)pyrazin-2-yl]-N-tert-butoxycarbonyl-carbamate (50 mg, 0.07865 mmol) in CH2Cl2 (5.0 mL) at 0° C. was added acetic acid (5.196 mg, 4.920 μL, 0.08652 mmol) and sodium triacetoxyborohydride (28.34 mg, 0.1337 mmol) and the resulting solution stirred at 0-10° C. for 2 h. The reaction mixture was diluted with saturated aqueous sodium hydrogen carbonate solution (5 mL) and dichloromethane (10 mL) and the layers separated. The aqueous layer was extracted further with dichloromethane (2×5 mL) and the combined organic extracts were then dried over MgSO 4 and concentrated in vacuo. The residue was taken up in CH2Cl2 (3.0 mL) and TFA (44.83 mg, 30.29 μL, 0.3932 mmol) was added and the resulting solution stirred at rt overnight. The reaction mixture was concentrated in vacuo and purified by reverse phase preparative HPLC [Waters Sunfire C18, 100 Å column, 5 M, 19 mm×150 mm, (solvent A: 0.05% TFA in water; solvent B: 0.05% TFA in CH3CN) over 16 minutes at 20 mL/min] The product fractions were collected and lyopholised to leave the title compound as a yellow solid (30.5 mg, 58% yield); 1H NMR (400 MHz, DMSO) δ 1.19 (d, 6H), 1.40-1.59 (m, 3H), 1.79 (m, 2H), 1.96 (m, 1H), 2.88 (m, 1H), 3.24 masked signal, 3.47 (m, 1H), 6.37 (t, 1H), 6.79 (m, 2H), 7.16 (br s, 2H), 7.64 (s, 1H), 7.78 (m, 2H), 7.94 (m, 2H), 8.28 (br s, 1H), 8.38 (m, 2H), 8.52 (br s, 1H) and 8.94 (s, 1H) ppm; MS (ES+) 533.3 (M+H), (ES−) 531.2 (M−H)
Workup
- customthe layers separated
- extractionThe aqueous layer was extracted further with dichloromethane (2×5 mL)
- dry with materialthe combined organic extracts were then dried over MgSO 4
- concentrationconcentrated in vacuo
- stirringthe resulting solution stirred at rt overnight
- concentrationThe reaction mixture was concentrated in vacuo
- custompurified by reverse phase preparative HPLC [Waters Sunfire C18, 100 Å column, 5 M, 19 mm×150 mm, (solvent A: 0.05% TFA in water; solvent B: 0.05% TFA in CH3CN) over 16 minutes at 20 mL/min] The product fractions
- customwere collected
Sources & evidence
- ORD record
- ord-1f306b191b264b8b84f5fa533576fc99
- Source dataset
- uspto-grants-2014_12
- Patent number
- US08912198B2
Reaction notes
Loading…