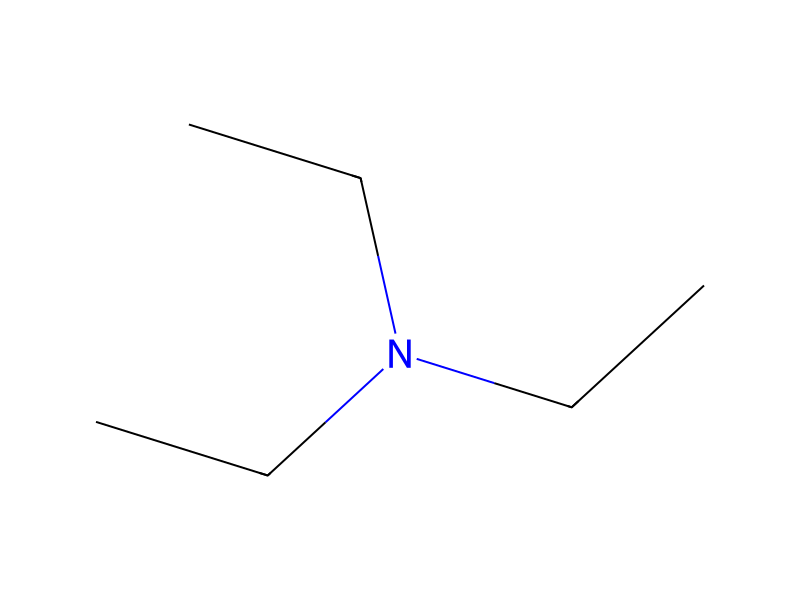
Triethylamine → HCID 15617222, compound
Reaction equation

[CH2:1]([O:3][C:4](=[O:17])[CH2:5][NH:6][C:7]1[C:12]([N+:13]([O-])=O)=[CH:11][CH:10]=[C:9]([Cl:16])[N:8]=1)[CH3:2].C(N(CC)CC)C.[Cl:25][C:26]1[CH:34]=[CH:33][C:29]([C:30](Cl)=[O:31])=[CH:28][CH:27]=1>O1CCCC1.[Pt](=O)=O>[CH2:1]([O:3][C:4](=[O:17])[CH2:5][NH:6][C:7]1[C:12]([NH:13][C:30](=[O:31])[C:29]2[CH:33]=[CH:34][C:26]([Cl:25])=[CH:27][CH:28]=2)=[CH:11][CH:10]=[C:9]([Cl:16])[N:8]=1)[CH3:2]Participants
Reactant 3


Product 1

Reagents, catalysts & solvents 2


Procedure
A solution of N-(6-chloro-3-nitro-2-pyridinyl) glycine ethyl ester (5.0 g, 0.0193 mole) in tetrahydrofuran (100 ml) was hydrogenated over platinum dioxide (0.5 g) at room temperature. After 2/3 of the theoretical amount of hydrogen had been absorbed, the mixture was filtered and a fresh portion of platinum dioxide (0.5 g) was added. The hydrogenation was continued until the theoretical amount of hydrogen was absorbed. The reaction mixture was cooled in ice, dried over magnesium sulfate and filtered. Triethylamine (2.05 g, 0.0203 mole) and 4-chlorobenzoyl chloride (3.55 g, 0.0203 mole) were added simultaneously, dropwise, to the cooled and stirred solution. After stirring at room temperature for 1.75 hours, the mixture was evaporated under reduced pressure and partitioned between water (100 ml) and ethyl acetate (100 ml). The layers were separated and the aqueous layer was re-extracted with ethyl acetate. The combined ethyl acetate layers were washed three times with saturated sodium chloride solution, dried over magnesium sulfate, charcoaled, filtered, and evaporated under reduced pressure. The resulting solid was dissolved in hot isopropyl alcohol, filtered hot and cooled to room temperature to give a solid which was collected by filtration and rinsed with isopropyl ether to give 2.2 g of title compound (31% yield). A 0.5 g portion was dissolved in hot isopropyl alcohol, filtered hot, isopropyl ether was added, and the mixture was cooled. The resulting suspension was filtered, the filtrate was evaporated under reduced pressure, and the residue was recrystallized from isopropyl alcohol/water and dried under high vacuum at 60° C. to give 0.37 g of the title compound, mp 157°-158.5° C.
Workup
- customAfter 2/3 of the theoretical amount of hydrogen had been absorbed
- filtrationthe mixture was filtered
- additiona fresh portion of platinum dioxide (0.5 g) was added
- customwas absorbed
- temperatureThe reaction mixture was cooled in ice
- dry with materialdried over magnesium sulfate
- filtrationfiltered
- temperaturedropwise, to the cooled
- stirringAfter stirring at room temperature for 1.75 hours
- customthe mixture was evaporated under reduced pressure
- custompartitioned between water (100 ml) and ethyl acetate (100 ml)
- customThe layers were separated
- extractionthe aqueous layer was re-extracted with ethyl acetate
- washThe combined ethyl acetate layers were washed three times with saturated sodium chloride solution
- dry with materialdried over magnesium sulfate
- filtrationfiltered
- customevaporated under reduced pressure
- dissolutionThe resulting solid was dissolved in hot isopropyl alcohol
- filtrationfiltered hot
- customto give a solid which
- filtrationwas collected by filtration
- washrinsed with isopropyl ether
Sources & evidence
- ORD record
- ord-3bb30af687f048019571987edf7e5ae7
- Source dataset
- uspto-grants-1990_08
- Patent number
- US04948800
Reaction notes
Loading…