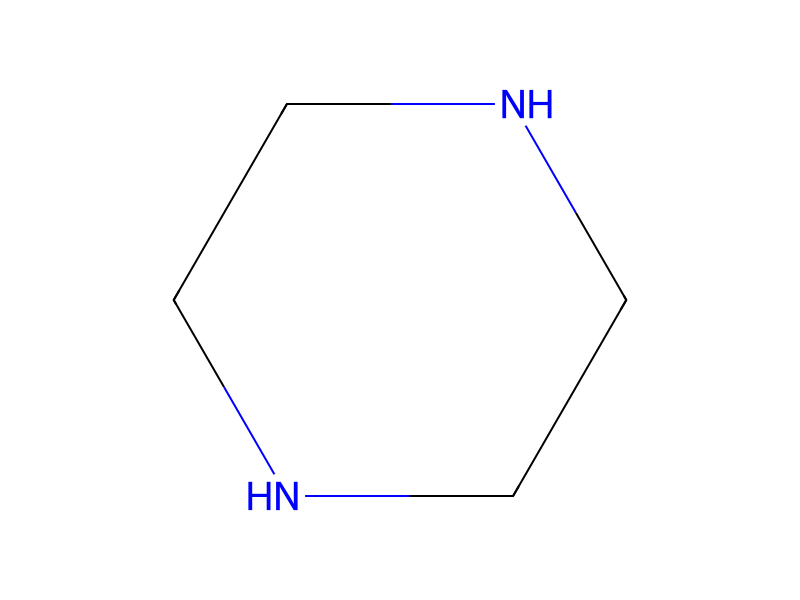
Piperazine → HCID 17723848, compound
Reaction equation

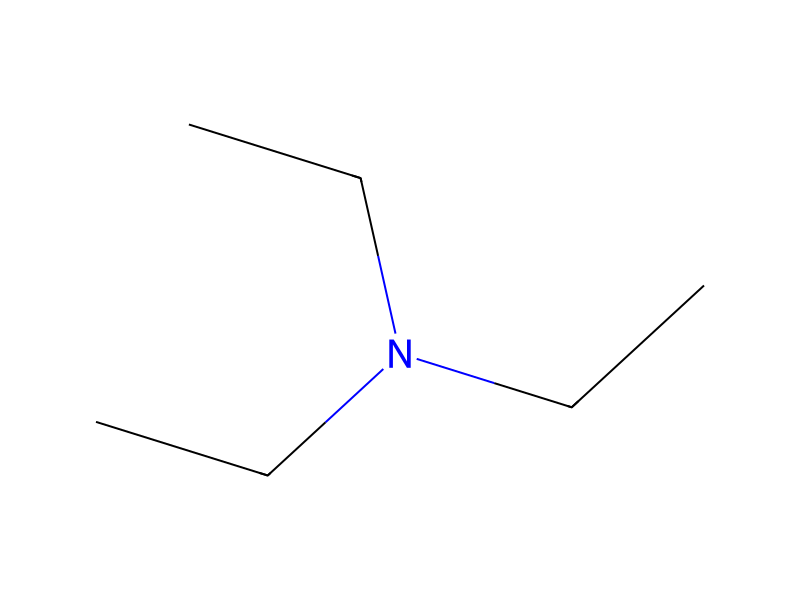
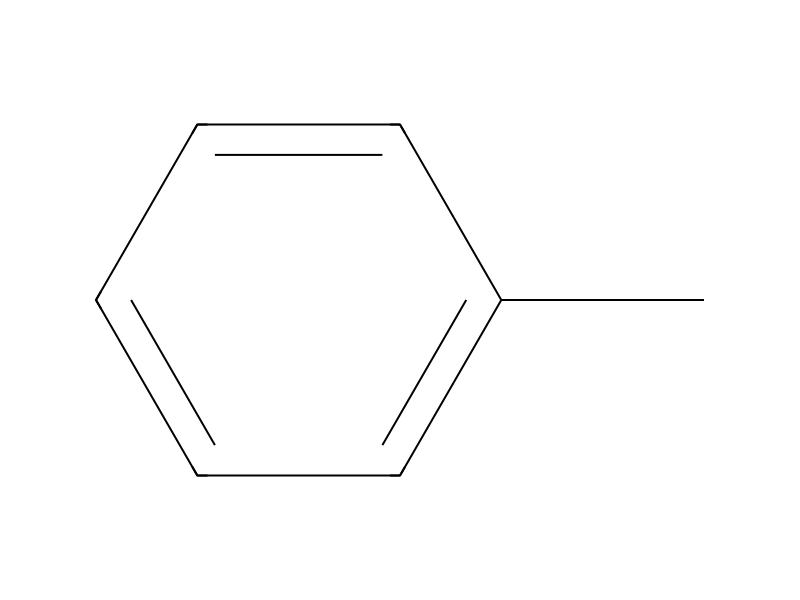
[NH:1]1[CH2:6][CH2:5][NH:4][CH2:3][CH2:2]1.[CH3:7][C:8]1[CH:15]=[CH:14][C:11]([CH2:12]Cl)=[CH:10][CH:9]=1.[OH-].[Na+].C(N(CC)CC)C.[C:25](OC([O-])=O)([O:27][C:28]([CH3:31])([CH3:30])[CH3:29])=[O:26]>C1(C)C=CC=CC=1.C(OCC)(=O)C>[CH3:7][C:8]1[CH:15]=[CH:14][C:11]([CH2:12][N:1]2[CH2:6][CH2:5][N:4]([C:25]([O:27][C:28]([CH3:31])([CH3:30])[CH3:29])=[O:26])[CH2:3][CH2:2]2)=[CH:10][CH:9]=1Participants



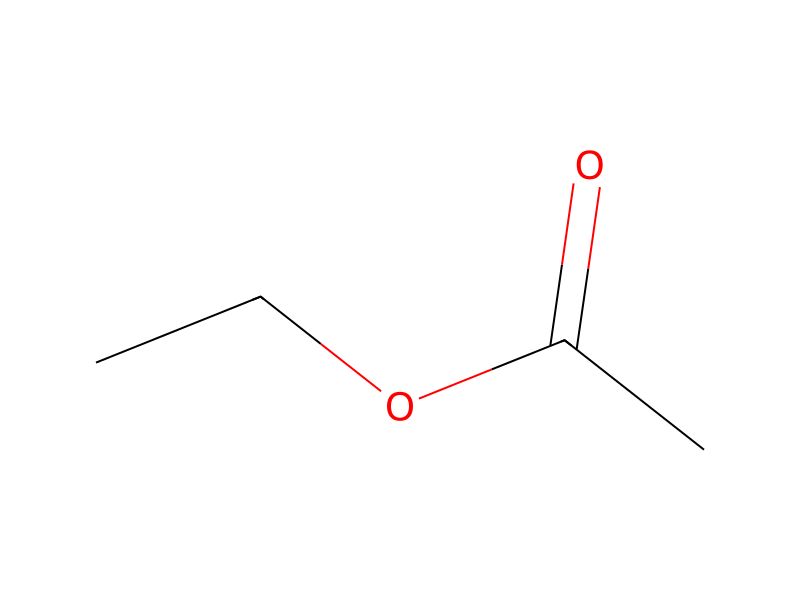
Conditions
- Temperature
- 85 °C
Procedure
Piperazine (170 g, 2.0 mol) was suspended in toluene (800 mL) and warmed to 85° C. The mixture was dissolved while stirring for 10 minutes. To the solution was added 4-methylbenzyl chloride (53 mL, 0.4 mol), and the mixture was stirred at 85° C. for 2 hours and then left to stand for cooling to 50° C. A 1 N aqueous sodium hydroxide solution (440 mL) and ethyl acetate (100 mL) were added thereto, and the process of phase separation was performed. The resulting organic layer was washed with a 1 N aqueous sodium hydroxide solution (400 mL) and dried over magnesium sulfate. The solvent was distilled off under reduced pressure, and the residue was dissolved in tetrahydrofuran (420 mL). Triethylamine (84 mL, 0.6 mol) was added thereto, and the resulting mixture was ice cooled. t-Butyl dicabonate (97 mL, 0.42 mol) was added thereto, and the mixture was stirred overnight while elevating the temperature to room temperature. The solvent was distilled off under reduced pressure. A 4 N aqueous sodium hydroxide solution (200 mL) and ethyl acetate (300 mL) were added thereto, and the process of phase separation was performed. The resulting organic layer was washed with an aqueous saturated sodium hydrogen carbonate solution (400 mL) and dried over magnesium sulfate. The solvent was distilled off under reduced pressure. To the residue was added a small amount of hexane, and the mixture was ice cooled. The obtained crystals were collected by filtration and washed with ice-cooled hexane to obtain the title compound (70 g, yield 60%). The mother liquor was concentrated under reduced pressure, and then the residue was purified with silica gel column chromatography (hexane to hexane:ethyl acetate=3:1). The resulting oily matter was left to stand at room temperature. The obtained crystals were collected by filtration and washed with ice-cooled hexane to obtain the title compound (22 g, yield 19%).
Workup
- dissolutionThe mixture was dissolved
- stirringthe mixture was stirred at 85° C. for 2 hours
- waitleft
- temperaturefor cooling to 50° C
- customthe process of phase separation
- washThe resulting organic layer was washed with a 1 N aqueous sodium hydroxide solution (400 mL)
- dry with materialdried over magnesium sulfate
- distillationThe solvent was distilled off under reduced pressure
- dissolutionthe residue was dissolved in tetrahydrofuran (420 mL)
- temperaturecooled
- stirringthe mixture was stirred overnight
- customto room temperature
- distillationThe solvent was distilled off under reduced pressure
- additionA 4 N aqueous sodium hydroxide solution (200 mL) and ethyl acetate (300 mL) were added
- customthe process of phase separation
- washThe resulting organic layer was washed with an aqueous saturated sodium hydrogen carbonate solution (400 mL)
- dry with materialdried over magnesium sulfate
- distillationThe solvent was distilled off under reduced pressure
- additionTo the residue was added a small amount of hexane
- temperaturecooled
- filtrationThe obtained crystals were collected by filtration
- washwashed with ice-
- temperaturecooled hexane
Sources & evidence
- ORD record
- ord-0951d3c40dae4426afac1246596ecf4e
- Source dataset
- uspto-grants-2010_09
- Patent number
- US07795267B2
Reaction notes
Loading…