Tetrahydrofuran → HCID 57712919, compound
Reaction equation

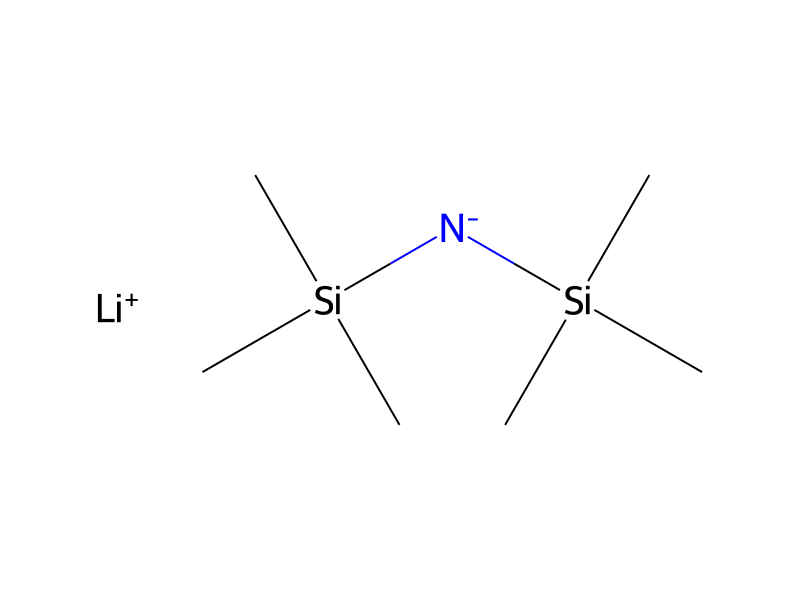
[Br-].[C:2]([CH2:5][CH2:6][CH2:7][P+](C1C=CC=CC=1)(C1C=CC=CC=1)C1C=CC=CC=1)([OH:4])=[O:3].C[Si]([N-][Si](C)(C)C)(C)C.[Li+].CO[C:39]1[C:40]([O:47][CH3:48])=[C:41]([CH:44]=[CH:45][CH:46]=1)[CH:42]=O.C1C[O:52][CH2:51]C1>>[CH3:48][O:47][C:40]1[CH:39]=[C:46]([O:52][CH3:51])[CH:45]=[CH:44][C:41]=1/[CH:42]=[CH:7]/[CH2:6][CH2:5][C:2]([OH:4])=[O:3]Participants




Conditions
- Temperature
- -15 °C
Procedure
An oven-dried 2 L 3-neck flask was secured over a dry ice/isopropanol bath and magnetic stirring plate, fitted with an oven-dried 500 ml dropping funnel, a magnetic stir bar, an argon inlet, and a thermometer. The flask was flushed with argon, charged with 71.3 grams (166 mmol) of 3-carboxypropyltriphenylphosphonium bromide followed by 400 ml of THF, and closed. An argon balloon was used to cap the addition funnel and the reaction mixture was stirred for a half hour while cooling. A mixture of 63.9 grams (381.8 mmol) of lithium bis (trimethylsilyl) amide in 400 ml of THF was added to the reaction mixture via an addition funnel, with stirring, after the temperature had gone below −50° C. and at such a rate as to keep the temperature at or below −40° C. When the addition was complete, the dry ice bath was carefully removed and replaced with a ice/salt (NaCl) bath to maintain a reaction temperature about −10 to −20° C. The reaction mixture was stirred for about 2 hours to give a clear orange mixture. The temperature was reduced to −40° C. with a dry ice bath and a solution of 27.6 grams (166 mmol) of dimethoxybenzaldehyde in about 320 ml of THF was added to the reaction mixture over a half hour via an addition funnel. The temperature was held at −18 to −20° C. with the ice/salt bath for two hours, then the flask was allowed to warm to room temperature. The reaction mixture was allowed to warm overnight, then transferred to a 3 L evaporation flask, and the solvents were evaporated. The by-product, triphenylphosphene oxide, and any remaining starting material was removed by extraction with a 1:1 mixture of ethyl acetate/water (1.4 L) and washing with saturated sodium bicarbonate (300 ml). The combined aqueous layers were washed with 600 ml of ether to remove residual triphenylphosphene oxide, and acidified with 10% citric acid solution until the pH of the mixture was around 2-3. This mixture, which contained some solids, was extracted with ethyl acetate (2×300 ml). The combined extracts were dried and evaporated in vacuo. The yield was 36.2 grams of solid product (>98%). 1H NMR (DMSO-d6) δ: 12.09 (s, 1H), 7.29 (d, J=8.2 Hz, 1H), 6.49 (m, 3H), 6.06 (m, 1H), 3.76 (s, 3H), 3.74 (s, 3H), 2.34 (br s, 4H).
Workup
- customAn oven-dried 2 L 3-neck flask was secured over a dry ice/isopropanol bath
- custommagnetic stirring plate, fitted with an oven-dried 500 ml
- customThe flask was flushed with argon
- customclosed
- customto cap the addition funnel
- temperaturewhile cooling
- additionwas added to the reaction mixture via an addition funnel
- stirringwith stirring
- customhad gone below −50° C.
- custombelow −40° C
- additionWhen the addition
- customthe dry ice bath was carefully removed
- customreplaced with a ice/salt (NaCl) bath
- stirringThe reaction mixture was stirred for about 2 hours
- customto give a clear orange mixture
- customThe temperature was reduced to −40° C. with a dry ice bath
- temperatureto warm to room temperature
- temperatureto warm overnight
- customtransferred to a 3 L evaporation flask
- customthe solvents were evaporated
- customwas removed by extraction with a 1:1 mixture of ethyl acetate/water (1.4 L)
- washwashing with saturated sodium bicarbonate (300 ml)
- washThe combined aqueous layers were washed with 600 ml of ether
- customto remove residual triphenylphosphene oxide
- extractionwas extracted with ethyl acetate (2×300 ml)
- customThe combined extracts were dried
- customevaporated in vacuo
Sources & evidence
- ORD record
- ord-b8d7703cc321448198da74935e0c5401
- Source dataset
- uspto-grants-2009_06
- Patent number
- US07541454B2
Reaction notes
Loading…