Citric Acid → HCID 86684900, compound
Reaction equation

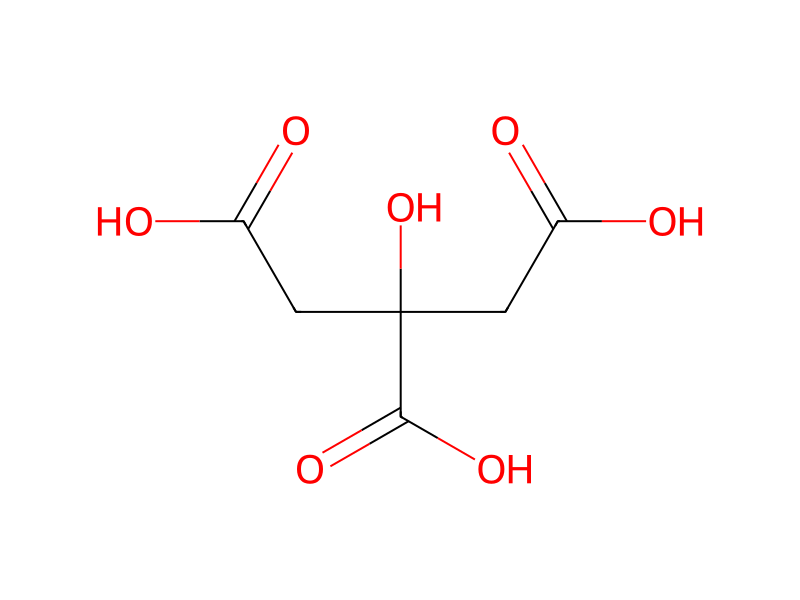
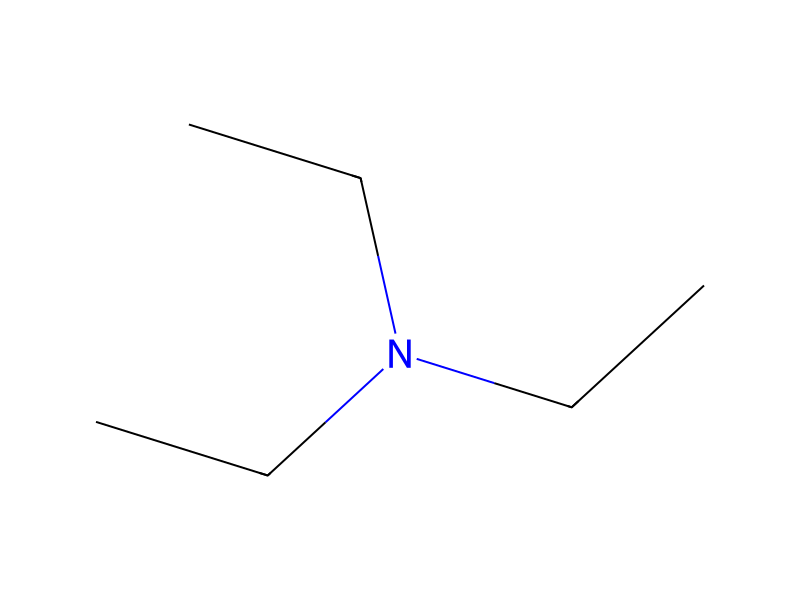
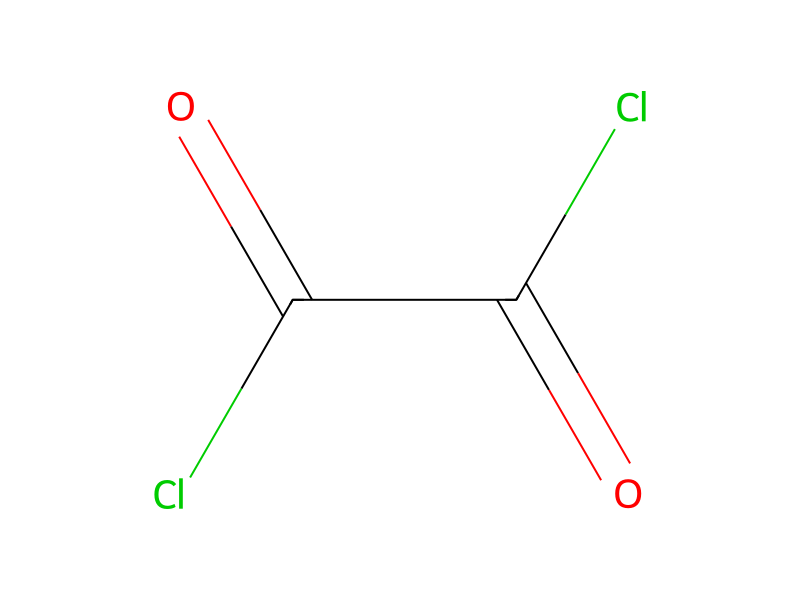
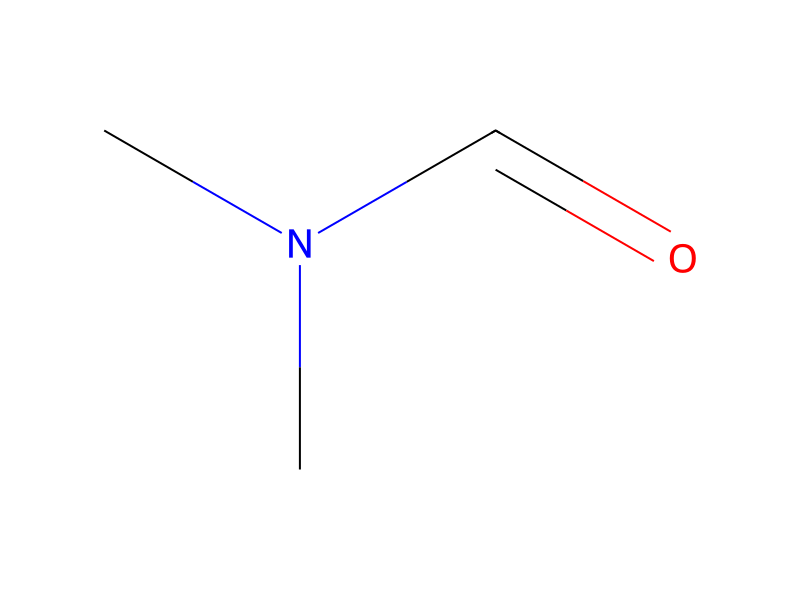
[CH3:1][C:2]1([C:5]([OH:7])=O)[CH2:4][CH2:3]1.C(Cl)(=O)C(Cl)=O.CS([O-])(=O)=O.[O:19]=[C:20]1[C@@H:25]2[CH2:26][C@@H:22]([CH2:23][NH2+:24]2)[O:21]1.C(N(CC)CC)C.C(O)(=O)CC(CC(O)=O)(C(O)=O)O>ClCCl.O.CN(C)C=O>[CH3:1][C:2]1([C:5]([N:24]2[CH2:23][C@@H:22]3[CH2:26][C@H:25]2[C:20](=[O:19])[O:21]3)=[O:7])[CH2:4][CH2:3]1Participants
Reactant 5


Product 1

Reagents, catalysts & solvents 3


Conditions
- Temperature
- 2 °C
Procedure
Methyl-cyclopropanecarboxylic acid (56.4 g, 552 mmol) was dissolved in dichloromethane (365 mL) and dimethylformamide (405 μl, 5.2 mmol) was added. The mixture was cooled to 2° C. and oxalyl chloride (70.8 g, 547 mmol) was added dropwise. It was allowed to warm and stirred for 90 min at room temperature. After that, it was added to a suspension of (1S,4S)-3-oxo-2-oxa-5-azonia-bicyclo[2.2.1]heptane methanesulfonate (110 g, 526 mmol) in dichloromethane (400 mL). The resulting suspension was cooled to 2° C. and triethylamine (256 mL, 1.84 mol) was added slowly (exothermic). After stirring for 70 min at room temperature, a solution of citric acid (81.0 g, 421 mmol) in water (550 mL) was added at 2° C. After separation of the phases, the aqueous phase was extracted with dichloromethane (300 mL). The combined organic extracts were washed with water (400 mL) and concentrated in vacuo to a volume of ca. 500 mL. Ethyl acetate (330 mL) was added and the residual dichloromethane was distilled off in vacuo (internal temperature 40° C.). Further ethyl acetate (50 mL) was added, internal temperature was increased to 50° C. and heptane (800 mL) was added slowly. Crystallization started after addition of ca. 300 mL heptane. The suspension was stirred for 12 h at room temperature and filtered. The crystals were washed with cold heptane (400 mL) and dried in vacuo at 40° C. to afford the title compound as colorless crystals (88.45 g, 86%). mp. 101-102° C. MS (ESI & APCI): m/z=196.1 [M+H]+. 1H NMR (CDCl3, 600 MHz); δ 0.60-0.67 (m, 2H), 0.87-0.91 (m, 1H), 1.13-1.17 (m, 1H), 1.39 (s, 3H), 2.04 (dd, J=1.2 Hz, 10.9 Hz, 1H), 2.32 (d, J=10.8 Hz, 1H), 3.59 and 3.69 (AB, JAB=11.5 Hz, each 1H), 4.97 (s, 1H), 5.18 (s, 1H).
Workup
- temperatureto warm
- temperatureThe resulting suspension was cooled to 2° C.
- stirringAfter stirring for 70 min at room temperature
- customAfter separation of the phases
- extractionthe aqueous phase was extracted with dichloromethane (300 mL)
- washThe combined organic extracts were washed with water (400 mL)
- concentrationconcentrated in vacuo to a volume of ca. 500 mL
- additionEthyl acetate (330 mL) was added
- distillationthe residual dichloromethane was distilled off in vacuo (internal temperature 40° C.)
- additionFurther ethyl acetate (50 mL) was added
- temperatureinternal temperature was increased to 50° C.
- additionheptane (800 mL) was added slowly
- customCrystallization
- additionafter addition of ca. 300 mL heptane
- stirringThe suspension was stirred for 12 h at room temperature
- filtrationfiltered
- washThe crystals were washed with cold heptane (400 mL)
- customdried in vacuo at 40° C.
Sources & evidence
- ORD record
- ord-129ead3dbd5c4e3087cdbf05a7d3729b
- Source dataset
- uspto-grants-2014_08
- Patent number
- US08796471B2
Reaction notes
Loading…