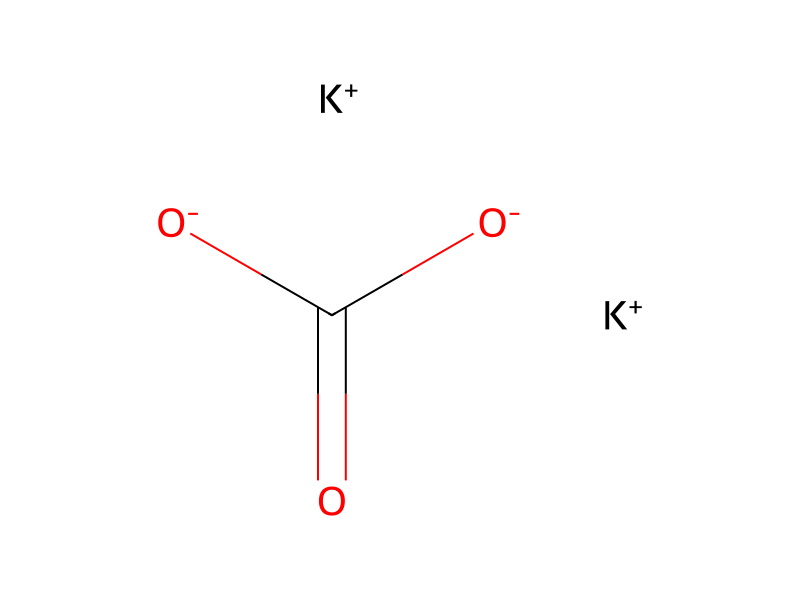
Potassium Carbonate → HCID 178463117, compound
Reaction equation

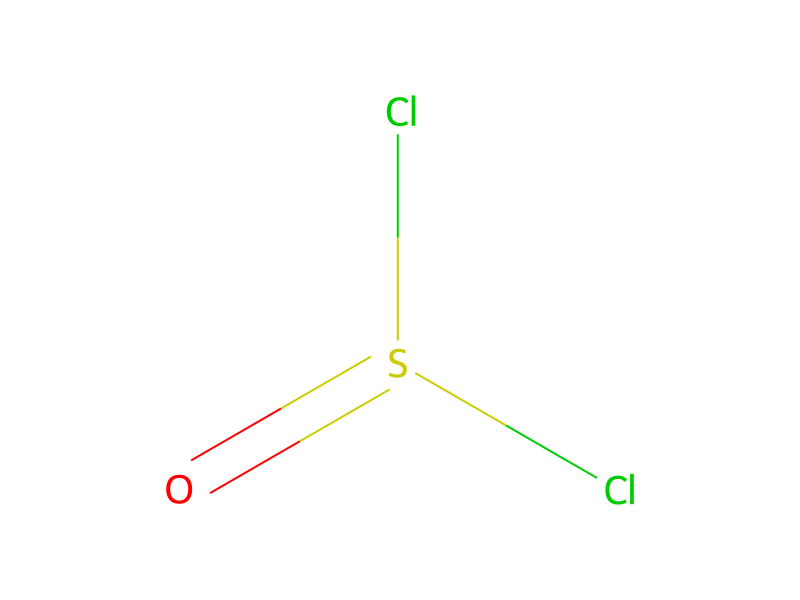
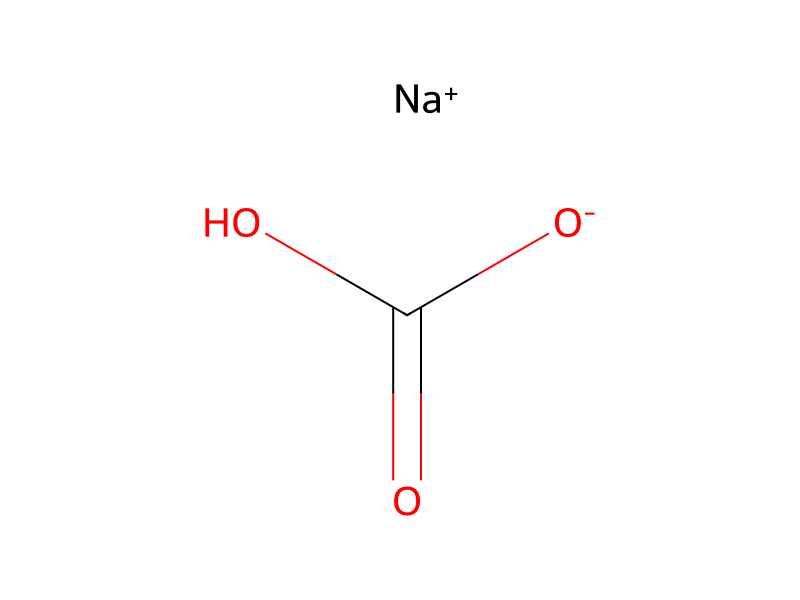
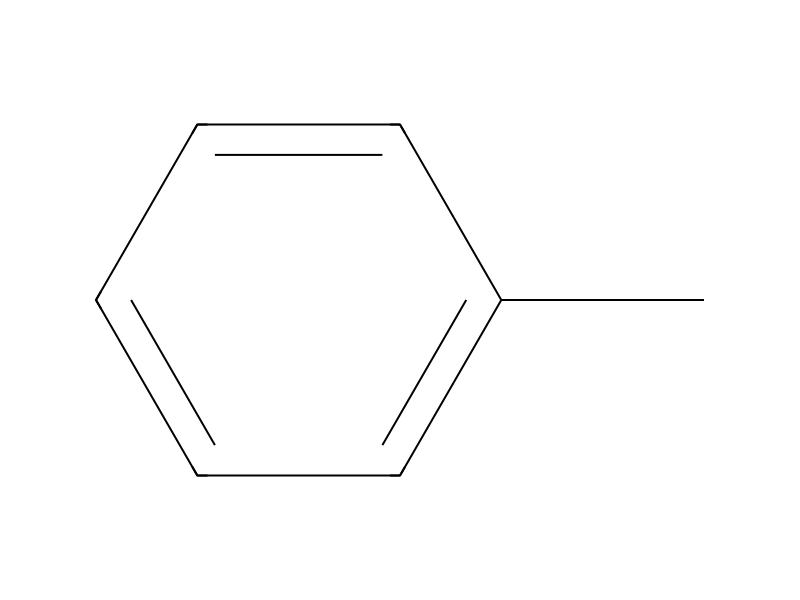
S(Cl)(Cl)=O.[Cl:5][C:6]1[C:14]([Cl:15])=[C:13]([S:16]([CH3:19])(=[O:18])=[O:17])[CH:12]=[CH:11][C:7]=1[C:8]([OH:10])=O.[CH2:20]([N:22]1[C:26]([OH:27])=[CH:25][CH:24]=[N:23]1)[CH3:21].C(=O)(O)[O-].[Na+].C(=O)([O-])[O-].[K+].[K+]>C1(C)C=CC=CC=1.[OH-].[Na+].O.CC(C)(O)C#N>[CH2:20]([N:22]1[C:26]([OH:27])=[C:25]([C:8](=[O:10])[C:7]2[CH:11]=[CH:12][C:13]([S:16]([CH3:19])(=[O:18])=[O:17])=[C:14]([Cl:15])[C:6]=2[Cl:5])[CH:24]=[N:23]1)[CH3:21]Participants
Reactant 5


Product 1

Reagents, catalysts & solvents 4


Conditions
- Temperature
- 90 °C
Procedure
Thionyl chloride (3.5 g, 6.7 mmol) was added to a slurry of 1.3 g (4.8 mmol) of 2,3-dichloro-4-methylsulfonylbenzoic acid in 10 mL of toluene with stirring and the mixture was heated at reflux until a homogeneous solution was obtained, which required about 45 min. The volatile components of the mixture were removed by evaporation under reduced pressure and the residual acid chloride was taken up in 10 mL of dichloromethane. This solution was combined with a solution of 5.4 g (4.8 mmol) of 1-ethyl-5-hydroxypyrazole dissolved in 6.2 mL of 1 N aqueous sodium hydroxide solution diluted with 5 mL of water with stirring at ambient temperature. After 1 hour, the pH was adjusted to 8 by adding 1 M aqueous sodium bicarbonate. The phases present were separated and the aqueous phase was extracted with 3 mL of dichloromethane. The organic phase and extract were combined, washed with 5 mL of water, dried over sodium sulfate, filtered, and concentrated by evaporation under reduced pressure. The residue was taken up in 15 mL of anhydrous dioxane and 730 mg (5.3 mmol) of solid potassium carbonate was added. The mixture was heated at 90° C. with stirring for 2 hours and then another 100 mg of potassium carbonate and 2-3 drops of acetone cyanohydrin were added. Heating and stirring were continued for another 30 min and then the mixture was cooled and poured into 15 mL of water. The phases were separated and the aqueous phase was extracted with 2×5 mL of dichloromethane. The aqueous phase was then acidified with 6 N aqueous hydrochloric acid and the solids that formed were extracted with 2×5 mL of dichloromethane. All of the organic phases and extracts were combined, washed with 5 mL of water, dried over sodium sulfate, filtered, and concentrated by evaporation under reduced pressure. The residue, which amounted to 1.4 g (80 percent of theory), was recrystallized from aqueous ethanol to obtain the title compound as a white solid melting at 198-200° C.
Workup
- temperaturethe mixture was heated
- temperatureat reflux until a homogeneous solution
- customwas obtained
- customabout 45 min
- customThe volatile components of the mixture were removed by evaporation under reduced pressure
- stirringwith stirring at ambient temperature
- customThe phases present were separated
- extractionthe aqueous phase was extracted with 3 mL of dichloromethane
- extractionThe organic phase and extract
- washwashed with 5 mL of water
- dry with materialdried over sodium sulfate
- filtrationfiltered
- concentrationconcentrated by evaporation under reduced pressure
- stirringwith stirring for 2 hours
- temperatureHeating
- stirringstirring
- waitwere continued for another 30 min
- temperaturethe mixture was cooled
- customThe phases were separated
- extractionthe aqueous phase was extracted with 2×5 mL of dichloromethane
- customthe solids that formed
- extractionwere extracted with 2×5 mL of dichloromethane
- washwashed with 5 mL of water
- dry with materialdried over sodium sulfate
- filtrationfiltered
- concentrationconcentrated by evaporation under reduced pressure
- customwas recrystallized from aqueous ethanol
Sources & evidence
- ORD record
- ord-0e0b69f55bba4c06858a52030a3f61b0
- Source dataset
- uspto-grants-2001_04
- Patent number
- US06211403B1
Reaction notes
Loading…