1,2-Epoxybutane → HCID 10182916, compound
Reaction equation

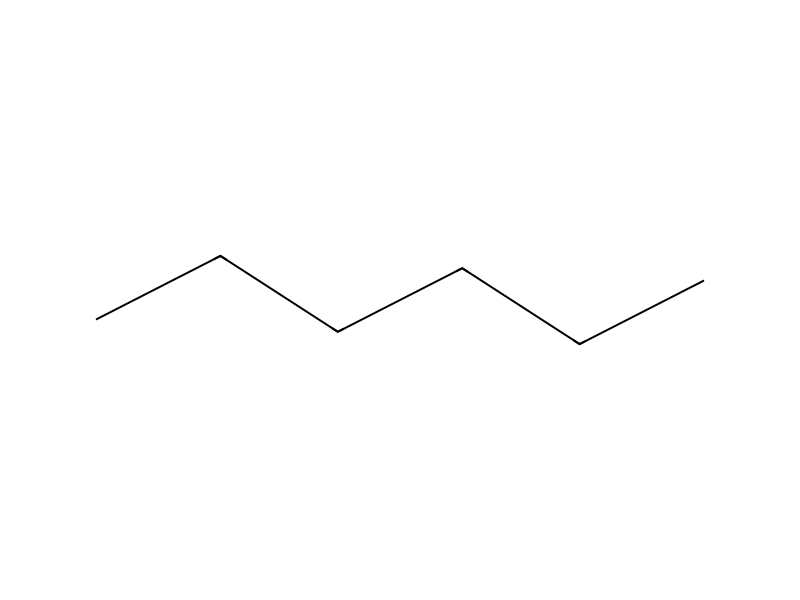
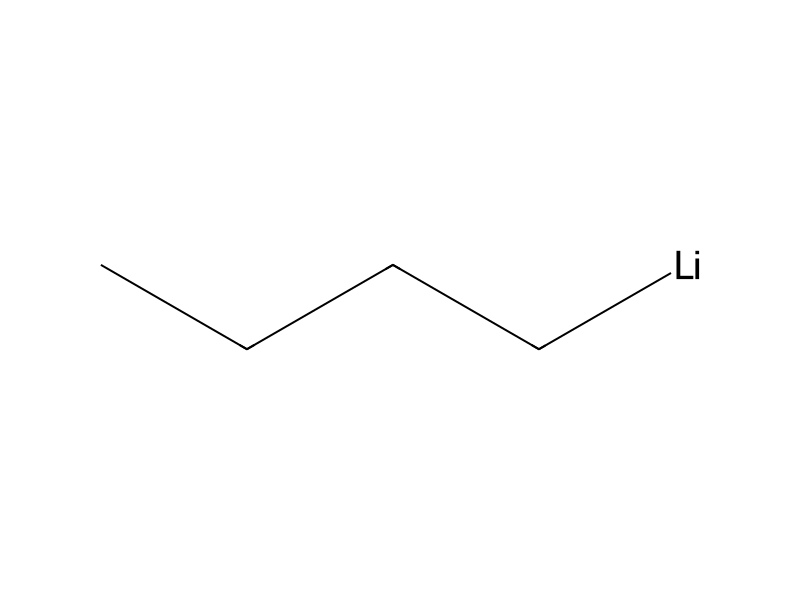
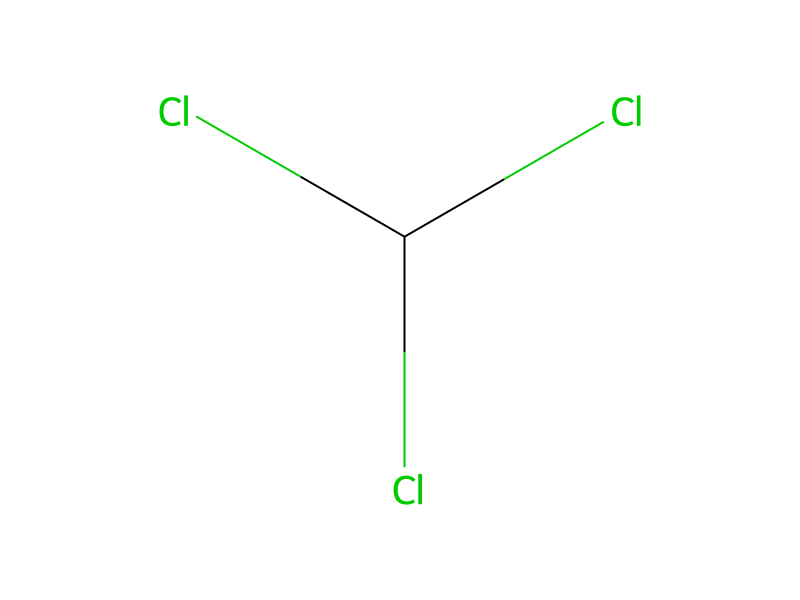
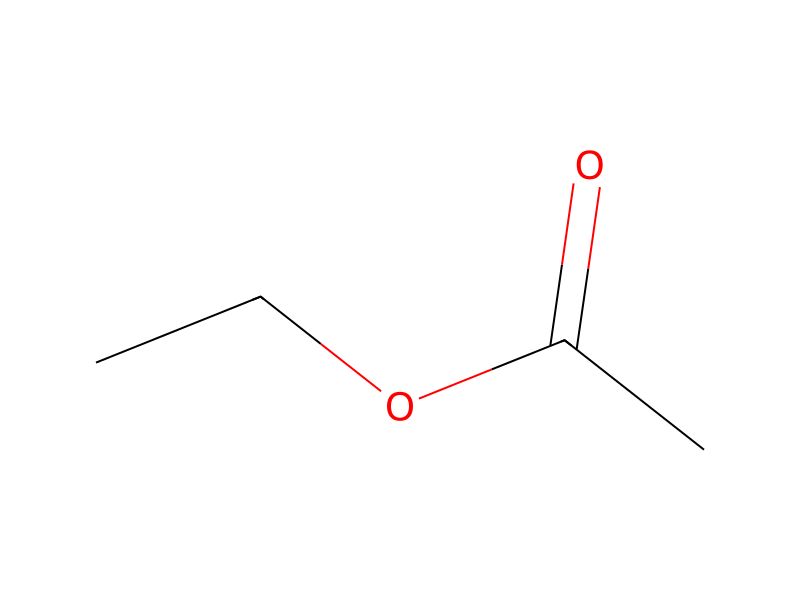
[OH:1][CH:2](CC)[CH2:3][C:4]1[C:5]([NH:16][C:17](=O)[C:18](C)([CH3:20])[CH3:19])=[CH:6][C:7]2[C:12]([C:13]=1[O:14][CH3:15])=[CH:11][CH:10]=[CH:9][CH:8]=2.COC1C2C(=CC=CC=2)C=C([NH:37][C:38](=[O:43])C(C)(C)C)C=1.C([Li])[CH2:45][CH2:46][CH3:47].CCC[CH2:52][CH2:53][CH3:54].[CH2:55]([CH:57]1[CH2:59][O:58]1)C.[O:60]1CCCC1>[Cl-].[NH4+].C(OCC)(=O)C.C(Cl)(Cl)Cl>[CH2:57]([O:58][C:59](=[O:60])[CH2:15][O:14][C:13]1[C:12]2[CH:11]=[CH:10][CH:9]=[CH:8][C:7]=2[CH:6]=[C:5]2[C:4]=1[C:3]([C:2](=[O:1])[C:38]([NH2:37])=[O:43])=[C:54]([CH2:53][CH3:52])[N:16]2[CH2:17][C:18]1[CH:20]=[CH:47][CH:46]=[CH:45][CH:19]=1)[CH3:55]Participants
Reactant 6




Product 1

Reagents, catalysts & solvents 3
Procedure
Commercially Available A. Preparation of N-(4-methoxynaphthalen-2-yl)-2,2-dimethylpropionamide. A solution of 3-amino-1-methoxynaphthalene (5.0 g, 29 mmol) in acetone (200 mL) was treated with pivaloyl chloride (4.0 mL, 33 mmol) and the resulting suspension stirred at room temperature for 1 h. Excess sodium bicarbonate was added and the resulting mixture stirred for 1 h, filtered, and reduced in vacuo to a volume of approximately 100 mL. The mixture was diluted with water and the resulting precipitate collected via vacuum filtration. The solid was washed with water and dried under vacuum for 60° C. to provide the title compound in quantitative yield: mp 169-170° C. 1H NMR (CDCl3) δ8.14 (d, J=7.7 Hz, 1H), 7.67 (d, J=7.1 Hz, 1H), 7.50 (s, 1H), 7.48 (bs, 1H), 7.43 (t, J=7.0 Hz, 1H), 7.35 (t, J=7.0 Hz, 1H), 7.23 (s, 1H), 4.00 (s, 3H), 1.35 (s, 9H); MS FD+ m/e 257 (p); IR (CHCl3, cm−1) 3450, 3000, 1678, 1531. Anal. Calcd for C16H19NO2: C, 74.68; H, 7.44; N, 5.44. Found: C, 74.76; H, 7.68; N, 5.58. B. Preparation of N-[3-(2-hydroxybutyl)-4-methoxynaphthalen-2-yl]-2,2-dimethylpropionamide. To a solution of N-(4-methoxynaphthalen-2-yl)-2,2-dimethylpropionamide (514 mg, 2.00 mmol) in tetrahydrofuran (4 mL) cooled in an ice/acetone bath was added 2.0 M n-butyllithium in hexane (3.0 mL, 6.0 mmol) slowly via syringe. The mixture was allowed to warm to room temperature over 30 min and an additional portion of tetrahydrofuran (5 mL) was added. 2-Ethyloxirane (0.57 mL, 6.6 mmol) was added and the resulting mixture was stirred at room temperature for 40 h. The mixture was diluted with aqueous saturated ammonium chloride solution and ethyl acetate. The organic layer was washed once with saturated sodium bicarbonate solution, separated, dried (sodium sulfate), filtered, and concentrated in vacuo. Chromatography (silica gel, methylene chloride/ethyl acetate) of the residue provided 396 mg (60%) of the title compound and 140 mg (27%) of starting material: mp 173-174° C. 1H NMR (CDCl3) δ9.19 (bs, 1H), 8.18 (s, 1H), 7.97 (m, 1H), 7.80 (m, 1H), 7.42 (m, 2H), 3.93 (m, 1H), 3.90 (s, 3H), 3.15 (dd, J=13.9, 2.0 Hz, 1H), 2.73 (dd, J=14.3, 9.4 Hz, 1H), 2.05 (bs, 1H), 1.76 (septet, J=7.5 Hz, 1H), 1.65 (septet, J=7.2 Hz, 1H), 1.36 (s, 9H), 1.07 (t, J=7.5 Hz, 3H); MS ES− m/e 328 (p−1). IR (CHCl3, cm−1) 3400 (b), 3009, 1667, 1531. Anal. Calcd for C20H27NO3: C, 72.92; H, 8.26; N, 4.25. Found: C, 72.72; H, 8.29; N, 4.40. C. Preparation of 2,2,2-trifluoro-N-[3-(2-hydroxybutyl)-4-methoxynaphthalen-2-yl]acetamide. To a mixture N-[3-(2-hydroxybutyl)-4-methoxynaphthalen-2-yl]-2,2-dimethylpropionamide (3.57 g, 10.8 mmol) and lithium hydroxide (0.52 g, 22 mmol) in methyl sulfoxide (8 mL) was added 1M aqueous lithium hydroxide solution (2 mL) and a trace of methanol. The mixture was warmed to 180° C. while stirring and allowing some volatile material to distill away. After 1 h toluene (2 mL) was added and the resulting mixture stirred for 3 h and allowed to cool to room temperature. A solid mass formed that was broken up by the addition of water and ethyl acetate. The mixture was saturated with sodium chloride and the layers separated. The aqueous layer was extracted with an additional portion of ethyl acetate. The combined ethyl acetate extracts were washed once with water, dried (magnesium sulfate), filtered, and concentrated in vacuo. The residue was dissolved in methylene chloride (20 mL) and treated with pyridine (2.6 mL, 32 mmol) and trifluoroacetic anhydride (3.1 mL, 22 mmol). The mixture was stirred for 20 h and partitioned between ether and 10% aqueous citric acid. The aqueous layer was washed with an additional portion of ether. The combined ether extracts were washed once with saturated sodium bicarbonate solution, dried (magnesium sulfate), filtered, and concentrated in vacuo. The residue was dissolved in methanol and an excess of potassium carbonate was added. The mixture was stirred 30 min and partitioned between ether and water. The organic layer was separated, dried (magnesium sulfate), filtered, and concentrated in vacuo to reveal a brown oil which crystallized on standing to give 3.28 g (89%) of the title compound: mp 120-123° C. 1H NMR (CDCl3) δ10.63 (bs, 1H), 8.19 (s, 1H), 8.01 (m, 1H), 7.82 (m, 1H), 7.48 (m, 2H), 3.95 (m, 1H), 3.90 (s, 3H), 3.22 (dd, J=14.3, 2.2 Hz, 1H), 2.77 (dd, J=14.3, 8.8 Hz, 1H), 2.23 (bs, 1H), 1.76 (septet, J=7.0 Hz, 1H), 1.58 (septet, J=7.0 Hz, 1H), 1.04 (t, J=7.3 Hz, 3H); MS ES− m/e 341 (p). IR (CHCl3, cm−1) 3465 (b), 1701, 1576, 1182. Anal. Calcd for C17H18F3NO3: C, 59.82; H, 5.32; N, 4.10. Found: C, 59.64; H, 5.13; N, 4.02. D. Preparation of 2,2,2-trifluoro-N-[4-methoxy-3-(2-oxobutyl)-naphthalen-2-yl]acetamide. To a solution of 2,2,2-trifluoro-N-[3-(2-hydroxybutyl)-4-methoxynaphthalen-2-yl]acetamide (6.24 g, 18.3 mmol) in methylene chloride (100 mL) was added pyridinium chlorochromate (6.20 g, 28.8 mmol) and the resulting mixture stirred at room temperature for 24 h. Silica gel (25 g) was added and the resulting suspension stirred for 1 h. The mixture was filtered through a pad of silica gel that was washed thoroughly with methylene chloride. The filtrate was concentrated in vacuo and the residue dissolved in a minimum of methylene chloride. This solution was diluted with hexane and the resulting precipitate collected via vacuum filtration. Cooling of the filtrate provided additional material to provide a total of 4.03 g (65%) of the title compound as yellow needles: mp 130-131° C. 1H NMR (CDCl3) δ10.08 (bs, 1H), 8.19 (s, 1H), 8.01 (m, 1H), 7.83 (m, 1H), 7.50 (m, 2H), 3.96 (s, 2H), 3.93 (s, 3H), 2.78 (q, J=7.3 Hz, 2H), 1.07 (t, J=7.3 Hz, 3H); MS ES− m/e 338 (p−1). IR (KBr, cm−1) 3268, 1706, 1166. Anal. Calcd for C17H16F3NO3: C, 60.18; H, 4.75; N, 4.13. Found: C, 59.92; H, 4.48; N, 4.05. E. Preparation of 2-ethyl-4-methoxy-1H-benz[f]indole. A solution of 2,2,2-trifluoro-N-[4-methoxy-3-(2-oxobutyl)-naphthalen-2-yl]acetamide (173 mg, 0.510 mmol) in methanol (3 mL) was treated with 1 M aqueous lithium hydroxide solution (1 mL) and heated at 95° C. for 4 h and 60° C. for 18 h. The mixture was cooled to room temperature and diluted with ether. The ether layer was washed once with water and the aqueous layer separated and extracted with a fresh portion of ether. The combined ether extracts were washed once with water, dried (sodium sulfate), filtered and concentrated in vacuo. Chromatography (silica gel, methylene chloride/hexane) of the residue provided 94 mg (82%) of the title compound as an oil. 1H NMR (CDCl3) δ8.25 (d, J=7.3 Hz, 1H), 7.82 (d, J=7.3 Hz, 1H), 7.77 (bs, 1H), 7.43 (s, 1H), 7.31 (m, 2H), 6.47 (s, 1H), 4.20 (s, 3H), 2.83 (q, J=7.7 Hz, 2H), 1.39 (t, J=7.3 Hz, 3H); MS ES− m/e 226 (p+1). IR (CHCl3, cm−1) 3472, 3009, 1418, 1314. Anal. Calcd for C15H15NO: C, 79.97; H, 6.71; N, 6.22. Found: C, 79.84; H, 6.82; N, 6.12. F. Preparation of 1-benzyl-2-ethyl-4-methoxy-1H-benz[f]indole. To a solution of 2-ethyl-4-methoxy-1H-benz[f]indole (548 mg, 2.43 mmol) in N,N-dimethylformamide (5 mL) was added benzyl bromide (0.64 mL, 5.8 mmol) followed by a 60% suspension of sodium hydride in mineral oil (290 mg, 7.3 mmol). After stirring for 1 h at room temperature, the mixture was diluted with water and extracted twice with ethyl acetate. The combined organic layers were washed three times with water, dried (magnesium sulfate), filtered, and concentrated in vacuo. Chromatography (silica gel, methylene chloride/hexanes) of the residue provided 635 mg of the title compound. MS ES+ m/e 316 (p+1) G. Preparation of 1-benzyl-2-ethyl-1,9-dihydrobenzo[f]indol-4-one. To a solution of 1-benzyl-2-ethyl-4-methoxy-1H-benz[f]indole (698 mg, 2.21 mmol) in methylene chloride (5 mL) was cooled to −10° C. and treated with boron tribromide (0.47 mL, 5.0 mmol). The mixture was warmed to 10° C. over 2.5 h then diluted with methanol. The mixture was diluted with methylene chloride and the resulting solution washed once with saturated sodium bicarbonate solution. The aqueous layer was extracted with a fresh portion of methylene chloride. The combined organic layers were washed once with water, dried (sodium sulfate), filtered, and concentrated in vacuo. Chromatography (silica gel, hexane/methylene chloride to acetone/methylene chloride) provided 310 mg (46%) of the title compound. 1H NMR (CDCl3) δ8.38 (d, J=7.3 Hz, 1H), 7.44 (m, 2H), 7.30 (m, 4 H), 6.94 (d, J=7.0 Hz, 2H), 6.66 (S, 1H), 5.17 (s, 2H), 3.95 (s, 2H), 2.52 (q, J=7.3 Hz, 2H), 1.26 (t, J=7.3 Hz, 3H); MS ES+ m/e 302 (p+1). IR (KBr, cm−1) 2962, 1618, 1290. Anal. Calcd for C21H19NO: C, 83.69; H, 6.35; N, 4.65. Found: C, 82.70; H, 6.40; N, 4.78. H. Preparation of (1-benzyl-2-ethyl-1H-benzo[f]indol-4-yloxy)acetic acid ethyl ester. A mixture of 1-benzyl-2-ethyl-1,9-dihydrobenzo[f]indol-4-one (301 mg, 1.00 mmol), cesium carbonate (325 mg, 1.00 mmol), and ethyl bromoacetate (0.17 mL, 15 mmol) in N,N-dimethylformamide (2 mL) was stirred at room temperature for 20 h. The mixture was diluted with ether and the resulting solution washed twice with water. The organic layer was separated, dried (magnesium sulfate), filtered, and concentrated in vacuo. Chromatography (silica gel, hexane/methylene chloride to acetone/methylene chloride) of the residue provided 168 mg (48%) of the title compound. 1H NMR (CDCl3) δ8.38 (d, J=8.8 Hz, 1H), 7.78 (d, J=9.5 Hz, 1H), 7.37 (s, 1H), 7.31 (m, 2H), 7.24 (m, 3H), 6.98 (d, J=8.1 Hz, 2H), 6.52 (s, 1H), 5.37 (s, 2H), 4.99 (s, 2H), 4.33 (q, J=7.0 Hz, 2H), 2.74 (q, J=7.3 Hz, 2H), 1.36 (t, J=7.3 Hz, 3H), 1.33 (t, J=7.0 Hz, 3H); MS ES+ m/e 388 (p+1). I. Preparation of 2-[[3-(2-amino-1,2-dioxoethyl)-1-benzyl-2-ethyl-1H-benz[f]indol-4-yl]oxy]acetic acid ethyl ester. A solution of (1-benzyl-2-ethyl-1H-benzo[f]indol-4-yloxy)acetic acid ethyl ester (168 mg, 0.434 mmol) in methylene chloride (10 mL) was cooled to −55° C. and treated with oxalyl chloride (1.0 mL, 11 mmol). The mixture was stirred for 2 h, warmed to −10° C., and concentrated in vacuo. The residue was dissolved in methylene chloride (10 mL) and treated with an excess of a solution of ammonia in dioxane. The mixture was stirred for 30 min and concentrated in vacuo. The residue was dissolved in methylene chloride which was washed once with water, dried (sodium sulfate), filtered, and concentrated in vacuo. HPLC purification of the residue provided 74 mg (37%) of the title compound as a yellow powder. 1H NMR (CDCl3) δ8.17 (d, J=7.7 Hz, 1H), 7.78 (d, J=6.8 Hz, 1H), 7.42 (s, 1H), 7.34 (m, 2H), 7.28 (m, 3H), 7.08 (d, J=8.1 Hz, 2H), 6.84 (bs, 1H), 5.49 (bs, 1H), 5.44 (s, 2H), 4.76 (s, 2H), 4.18 (q, J=6.9 Hz, 2H), 3.05 (q, J=7.3 Hz, 2H), 1.26 (t, J=7.7 Hz, 3H), 1.20 (t, J=7.3 Hz, 3H); MS ES+ m/e 459 (p+1).
Workup
- washThe organic layer was washed once with saturated sodium bicarbonate solution
- customseparated
- dry with materialdried (sodium sulfate)
- filtrationfiltered
- concentrationconcentrated in vacuo
Sources & evidence
- ORD record
- ord-1afd72c4655f47558182d107dfcabe9e
- Source dataset
- uspto-grants-2005_08
- Patent number
- US06930123B2
Reaction notes
Loading…