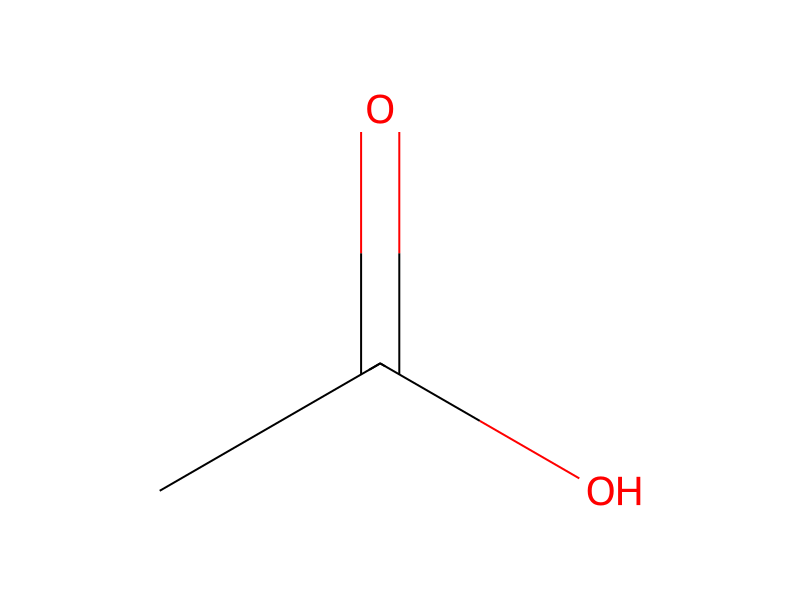
Acetic Acid → HCID 86707747, compound
Reaction equation

[C:1]1([C:7]2[CH:27]=[CH:26][C:10]3[N:11]=[C:12]([CH2:14][C:15]4[O:19][C:18]([CH2:20][NH:21][S:22](=[O:25])(=[O:24])[NH2:23])=[N:17][N:16]=4)[S:13][C:9]=3[CH:8]=2)[CH:6]=[CH:5][CH:4]=[CH:3][CH:2]=1.C[Si]([N-][Si](C)(C)C)(C)C.[Na+].C1COCC1.[CH3:43][S:44](Cl)(=[O:46])=[O:45].IC.[CH3:50]OC(C)(C)C.C(O)(=O)C>CN(C=O)C.CO>[CH3:43][S:44]([C:14]([C:15]1[O:19][C:18]([CH2:20][NH:21][S:22]([NH2:23])(=[O:24])=[O:25])=[N:17][N:16]=1)([C:12]1[S:13][C:9]2[CH:8]=[C:7]([C:1]3[CH:2]=[CH:3][CH:4]=[CH:5][CH:6]=3)[CH:27]=[CH:26][C:10]=2[N:11]=1)[CH3:50])(=[O:46])=[O:45]Participants
Reactant 7



Product 1

Reagents, catalysts & solvents 2
Procedure
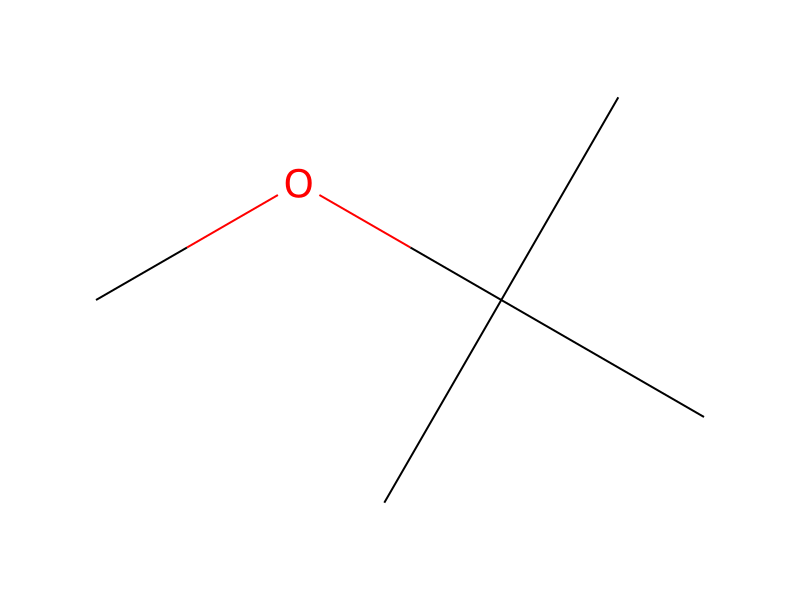
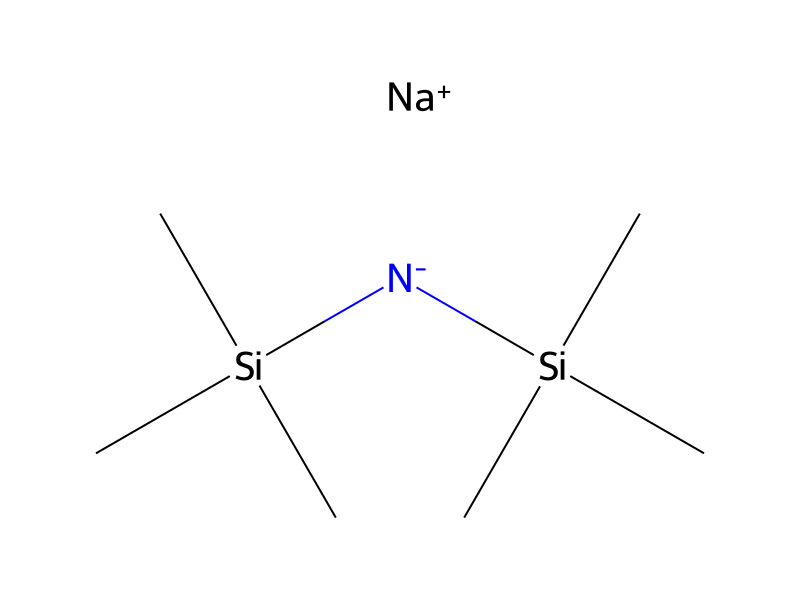
To a solution of Compound 46b (100 mg, 0.25 mmol) in DMF (3 mL) at 0° C. was added 1M NaHMDS in THF (0.55 mL, 0.55 mmol) and the mixture stirred for 10 min. Methanesulfonyl chloride (0.025 mL, 0.32 mmol) was added and the mixture stirred for 5 min then 2M iodomethane in tert-butyl methyl ether (0.22 mL, 0.45 mmol) was added and the reaction stirred for 1 h. Additional 2M iodomethane in tert-butyl methyl ether (0.080 mL, 0.16 mmol) was added and the reaction stirred for 1 h then quenched with acetic acid (0.043 mL, 0.75 mmol). The mixture was diluted with MeOH and purified by prep HPLC (Method B, gradient elution of 10-100% solvent B). Fractions containing product were made basic by the addition of 1.5 M phosphate buffer, evaporated under reduced pressure to remove most of the ACN, acidified by the addition of satd. NH4Cl and extracted with DCM (3×), dried (Na2SO4), filtered and concentrated under reduced pressure to afford the racemic product. The material was further purified by silica gel chromatography eluting with 0-100% EtOAc/hexane to afford the racemic product. The enantiomers were separated by chiral preparative chiral HPLC (Whelko 4.6×250 mm ID, 10 μm, 20 mL/min, 25% MeOH/EtOH (1:1) in heptane) to afford Reference 1, Isomer A (RT=14.3 min, 22 mg, 36% yield), LCMS=0.90 min using analytical method (M), 493.9 (M+H), 1H NMR (400 MHz, CDCl3) δ 8.05 (d, J=8.5 Hz, 1H), 8.01 (d, J=1.5 Hz, 1H), 7.68 (dd, J=8.8, 1.8 Hz, 1H), 7.56 (d, J=7.0 Hz, 2H), 7.47-7.40 (m, 2H), 7.39-7.31 (m, 1H), 5.73 (br. s., 2H), 4.67 (s, 2H), 3.20 (s, 3H), 2.44 (s, 3H), EL IC50=3751 nM; and Reference 1, Isomer B (RT=15.5 min, 23 mg, 37% yield), LCMS=0.90 min using analytical method (M), 493.9 (M+H), 1H NMR (400 MHz, CDCl3) δ 8.04 (d, J=8.8 Hz, 1H), 8.00 (d, J=1.3 Hz, 1H), 7.67 (dd, J=8.5, 1.5 Hz, 1H), 7.56 (d, J=7.0 Hz, 2H), 7.47-7.39 (m, 2H), 7.37-7.32 (m, 1H), 5.61 (br. s., 2H), 4.67 (s, 2H), 3.20 (s, 3H), 2.44 (s, 3H), EL IC50=5202 nM.
Workup
- stirringthe mixture stirred for 5 min
- stirringthe reaction stirred for 1 h
- stirringthe reaction stirred for 1 h
- custompurified by prep HPLC (Method B, gradient elution of 10-100% solvent B)
- additionFractions containing product
- additionwere made basic by the addition of 1.5 M phosphate buffer
- customevaporated under reduced pressure
- customto remove most of the ACN
- additionacidified by the addition of satd
- extractionNH4Cl and extracted with DCM (3×)
- dry with materialdried (Na2SO4)
- filtrationfiltered
- concentrationconcentrated under reduced pressure
- customto afford the racemic product
- customThe material was further purified by silica gel chromatography
- washeluting with 0-100% EtOAc/hexane
- customto afford the racemic product
- customThe enantiomers were separated by chiral preparative chiral HPLC (Whelko 4.6×250 mm ID, 10 μm, 20 mL/min, 25% MeOH/EtOH (1:1) in heptane)
- customto afford Reference 1, Isomer A (RT=14.3 min, 22 mg, 36% yield), LCMS=0.90 min
- customReference 1, Isomer B (RT=15.5 min, 23 mg, 37% yield), LCMS=0.90 min
Sources & evidence
- ORD record
- ord-898c4c288b0e440887db6e4e1b4feccf
- Source dataset
- uspto-grants-2014_03
- Patent number
- US08680090B2
Reaction notes
Loading…