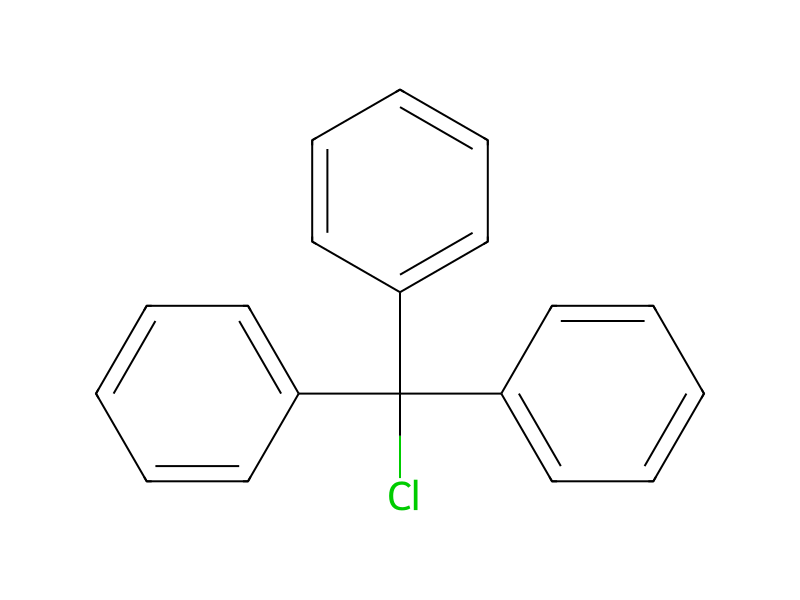
Trityl chloride → HCID 86758939, compound
Reaction equation

[OH:1][C@@H:2]1[CH2:21][C@@H:5]2[O:6][C:7]3[C:12]([CH2:13][CH2:14][P:15](=[O:20])([O:18][CH3:19])[O:16][CH3:17])=[CH:11][CH:10]=[CH:9][C:8]=3[C@@H:4]2[C@H:3]1[CH2:22][OH:23].C(N(CC)CC)C.C1(C(Cl)(C2C=CC=CC=2)C2C=CC=CC=2)C=CC=CC=1.[C:51](OC(=O)C)(=[O:53])[CH3:52].Cl.CO.C(=O)(O)[O-].[Na+]>CN(C=O)C.O>[C:51]([O:1][C@@H:2]1[CH2:21][C@@H:5]2[O:6][C:7]3[C:12]([CH2:13][CH2:14][P:15](=[O:20])([O:18][CH3:19])[O:16][CH3:17])=[CH:11][CH:10]=[CH:9][C:8]=3[C@@H:4]2[C@H:3]1[CH2:22][OH:23])(=[O:53])[CH3:52]Participants
Reactant 6


Product 1

Reagents, catalysts & solvents 2

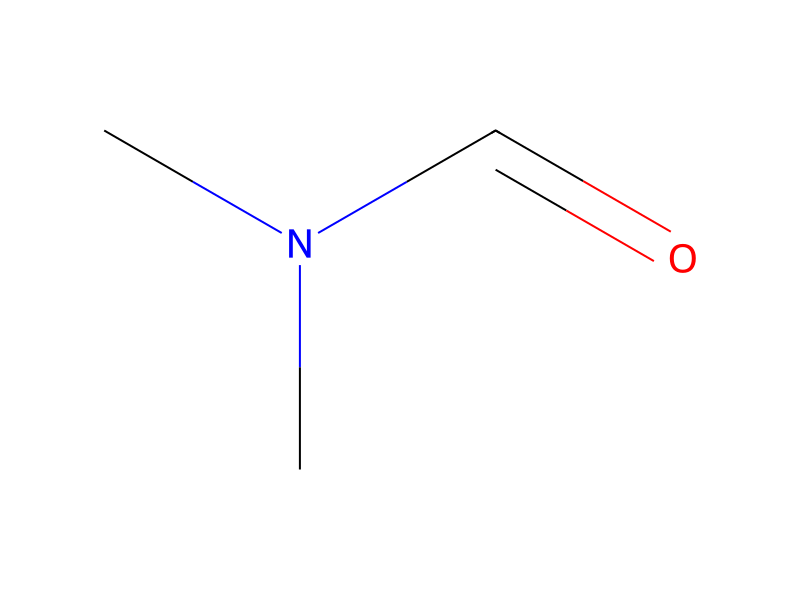
Procedure
A solution of dimethyl 2-{(1S,2R,3aS,8bS)-2,3,3a,8b-tetrahydro-2-hydroxy-1-hydroxymethyl-1H-cyclopenta[b]benzofuran-5-yl}ethylphosphonate (12) 1000 mg (2.921 mmol) and triethylamine 3.26 ml (23.368 mol) in anhydrous DMF (20 ml) was cooled to 0° C. under argon atmosphere, and a solution of triphenylmethyl chloride in DMF (5 ml×2) was added to the solution, followed by stirring at room temperature for 2 hours, and then at 60° C. overnight. The reaction mixture was again cooled to 0° C., and triethylamine 1.63 ml (11.684 mmol), N,N-dimethylamindpyridine 713.7 mg (5.842 mmol), and acetic anhydride 0.551 ml (5.842 mmol) were added to the reaction mixture. After the temperature was increased to 60° C., the mixture was stirred for 1 hour. After the mixture was allowed to cool to room temperature, ice-cold ion exchanged water 40 ml was added to the mixture, and the mixture was extracted with ethyl acetate (100 ml×3). The combined organic layer was washed with water (50 ml×2) and brine (50 ml), dried, concentrated, and then dried in vacuo. The thus-obtained yellow oil was dissolved in methanol 30 ml, and hydrogen chloride-methanol (1.387M) 6.32 ml (8.764 mmol) was added to the resultant solution, followed by stirring at room temperature for 2 hours. After sodium bicarbonate was added little by little to neutralize the mixture, the resultant suspension was filtered with Celite, and the filtrate was concentrated. After the precipitate was again filtered off with Celite, the filtrate was concentrated. The residue was subjected to column chromatography (Silica gel FL-60D produced by Fuji Silicia Co.; chloroform:methanol 95:5) to recover dimethyl 2-{(1S,2R,3aS,8bS)-2,3,3a,8b-tetrahydro-2-hydroxy-1-hydroxymethyl-1H-cyclopenta[b]benzofuran-5-yl}ethylphosphonate (12) used as the starting material (270.6 mg, 27%). The crude product of the titled compound was again purified by column chromatography (DIOL column produced by Yamazen Co.; ethyl acetate) to obtain the titled compound 667.9 mg (1.738 mmol) as a pale yellow oil (yield 60%). The structure of this compound was determined by the following data.
Workup
- customat 60° C.
- customovernight
- temperatureThe reaction mixture was again cooled to 0° C.
- temperatureAfter the temperature was increased to 60° C.
- stirringthe mixture was stirred for 1 hour
- temperatureto cool to room temperature
- additionwas added to the mixture
- extractionthe mixture was extracted with ethyl acetate (100 ml×3)
- washThe combined organic layer was washed with water (50 ml×2) and brine (50 ml)
- customdried
- concentrationconcentrated
- customdried in vacuo
- dissolutionThe thus-obtained yellow oil was dissolved in methanol 30 ml
- stirringby stirring at room temperature for 2 hours
- filtrationthe resultant suspension was filtered with Celite
- concentrationthe filtrate was concentrated
- filtrationAfter the precipitate was again filtered off with Celite
- concentrationthe filtrate was concentrated
- customThe residue was subjected to column chromatography (Silica gel FL-60D produced by Fuji Silicia Co.; chloroform:methanol 95:5)
- customto recover dimethyl 2-{(1S,2R,3aS,8bS)-2,3,3a,8b-tetrahydro-2-hydroxy-1-hydroxymethyl-1H-cyclopenta[b]benzofuran-5-yl}ethylphosphonate (12)
- customThe crude product of the titled compound was again purified by column chromatography (DIOL column produced by Yamazen Co.; ethyl acetate)
Sources & evidence
- ORD record
- ord-a6dabda5d79a4f5ba1b6ae2259bdde93
- Source dataset
- uspto-grants-2003_04
- Patent number
- US06555559B1
Reaction notes
Loading…