Iodine → HCID 66941079, compound
Reaction equation

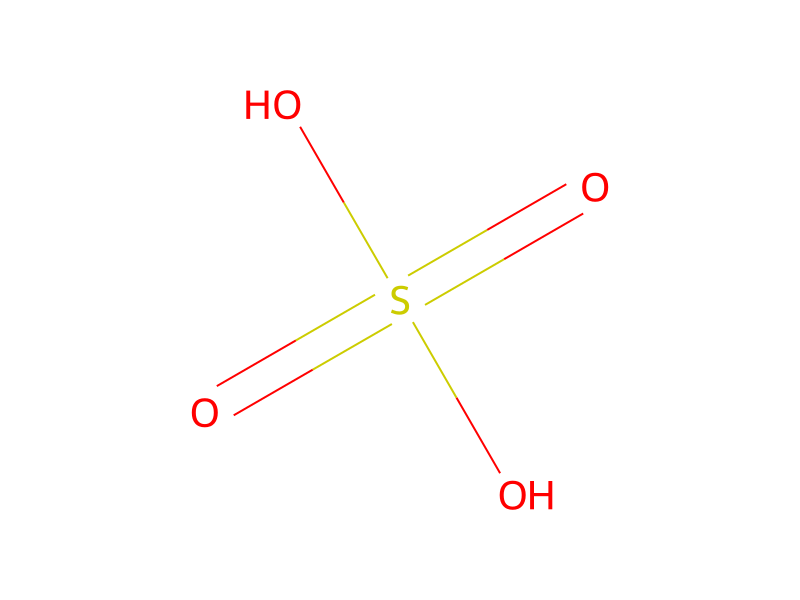
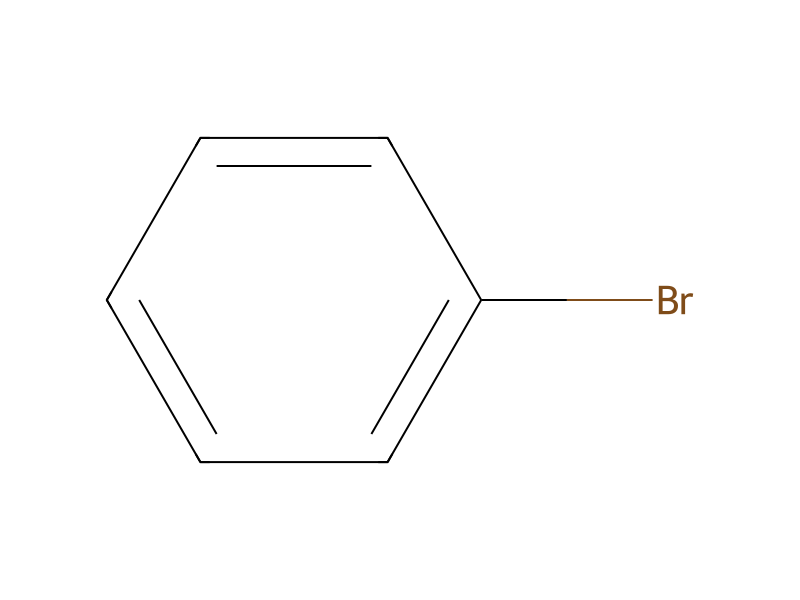
[Mg].II.Br[C:5]1[CH:10]=[CH:9][CH:8]=[CH:7][CH:6]=1.[CH3:11][CH:12]([CH2:28][CH2:29][CH2:30][CH:31]([CH3:33])[CH3:32])[CH2:13][CH2:14][O:15][C:16]1[CH:23]=[C:22]([CH:24]=[O:25])[C:21]([O:26][CH3:27])=[CH:20][C:17]=1[CH:18]=[O:19].C(=O)[C:35]1[CH:42]=[CH:41][C:38](C=O)=[CH:37][CH:36]=1.OS(O)(=O)=O>C1COCC1.C1(C)C=CC=CC=1>[C:5]1([CH:18]([OH:19])[C:17]2[CH:20]=[C:21]([O:26][CH3:27])[C:22]([CH:24]([C:35]3[CH:42]=[CH:41][CH:38]=[CH:37][CH:36]=3)[OH:25])=[CH:23][C:16]=2[O:15][CH2:14][CH2:13][CH:12]([CH3:11])[CH2:28][CH2:29][CH2:30][CH:31]([CH3:33])[CH3:32])[CH:10]=[CH:9][CH:8]=[CH:7][CH:6]=1Participants
Reactant 6


Product 1

Reagents, catalysts & solvents 2

Conditions
- Temperature
- 70 °C
Procedure
A 6 l four-neck flask with precision glass stirrer, reflux condenser and 2 dropping funnels was initially charged with 26.7 g (1.1 mol) of magnesium turnings. The apparatus was baked out under argon. At room temperature, 10 ml of absolute THF and a few crystals of iodine were added and the mixture was stirred briefly. Subsequently, a few ml of bromobenzene were added dropwise to the unstirred solution, and at the point of dropwise addition was heated briefly with a hot air blower. After the start of the reaction, a total (including the amount already added) of 215 g (1.37 mol) of bromobenzene were added dropwise within 45 min. At the same time, 490 ml of absolute THF were added. After the dropwise addition, the mixture was stirred under reflux for 1.5 h. Subsequently, a solution of 175.6 g (0.55 mol) of 2-(3,7-dimethyloctyloxy)-5-methoxyterephthalaldehyde in 600 ml of absolute THF were added dropwise with ice cooling. After half of the addition, the gel which had formed was admixed with 3000 ml of distilled toluene and dissolved with heating to 70° C. After cooling to room temperature, the remaining solution of terephthalaldehyde was added dropwise, and the viscous solution was stirred at 70° C. for a further 4 h. The reaction mixture was stirred into 4000 ml of ice-water with 40 ml of conc. H2SO4. The phases were separated and the aqueous phase was extracted with 1×500 ml of ethyl acetate. The combined organic phases were washed with 1×300 ml of water and dried over Na2SO4, and the solvent was removed under reduced pressure. The crude product was stirred with n-hexane twice at room temperature and once at 50° C., and filtered. Further purification was effected by melting the crude product at 65° C. in hexane. This operation was effected three times more. 157.1 g (60% of theory) of the product were obtained in 95% purity, which was used directly for the further synthesis. NMR (CDCl3): 7.38-7.44 (m, 4H), 7.24-7.35 (m, 6H), 7.09-9.12 (m, 1H), 7.04-7.07 (m, 1H), 6.54-6.57 (m, 2H, C(OH)H), 3.85-3.95 (m, 2H, OCH2), 3.76+3.77 (2×s, 3H, OCH3), 0.83-1.69 (m, 19H).
Workup
- temperatureA 6 l four-neck flask with precision glass stirrer, reflux condenser
- additionat the point of dropwise addition
- temperaturewas heated briefly with a hot air blower
- customAfter the start of the reaction
- additionwere added dropwise within 45 min
- additionAfter the dropwise addition
- stirringthe mixture was stirred
- temperatureunder reflux for 1.5 h
- additionAfter half of the addition
- customthe gel which had formed
- dissolutiondissolved
- temperatureAfter cooling to room temperature
- stirringthe viscous solution was stirred at 70° C. for a further 4 h
- customThe phases were separated
- extractionthe aqueous phase was extracted with 1×500 ml of ethyl acetate
- washThe combined organic phases were washed with 1×300 ml of water
- dry with materialdried over Na2SO4
- customthe solvent was removed under reduced pressure
- stirringThe crude product was stirred with n-hexane twice at room temperature and once at 50° C.
- filtrationfiltered
- customFurther purification
- customat 65° C.
- custom157.1 g (60% of theory) of the product were obtained in 95% purity, which
- customwas used directly for the further synthesis
Sources & evidence
- ORD record
- ord-a8c282fe35904120aac994a15a2e5c81
- Source dataset
- uspto-grants-2011_02
- Patent number
- US07884175B2
Reaction notes
Loading…