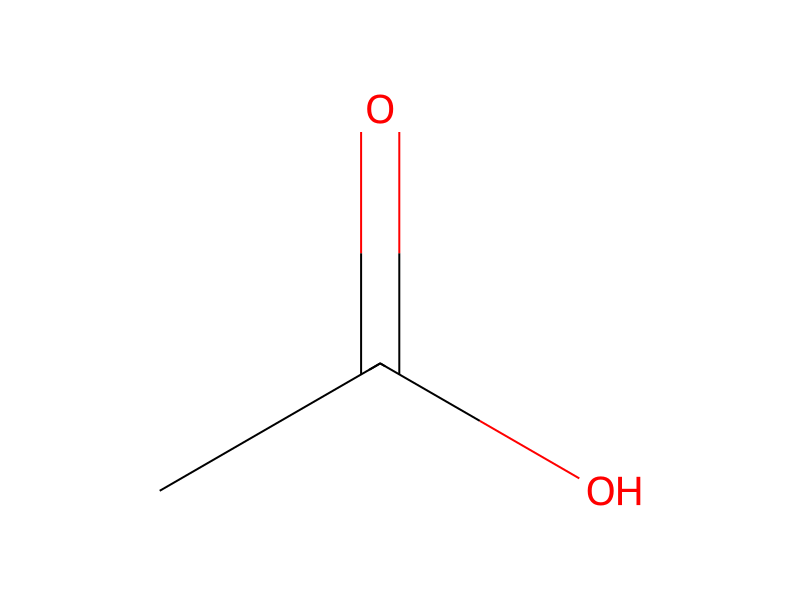
Acetic Acid → HCID 21702944, compound
Reaction equation

C([C@H]1COC(=O)N1C(=O)CC(C)(C)C(O[CH2:16][C:17]1[CH:22]=[CH:21][CH:20]=[CH:19][CH:18]=1)=O)(C)C.C[Si]([N-][Si](C)(C)C)(C)C.[K+].C(C1C=C(C(C)C)C=C(C(C)C)C=1S([N:54]=[N+:55]=[N-:56])(=O)=O)(C)C.[C:57]([OH:60])(=[O:59])[CH3:58].OO.[O-]S([O-])(=S)=O.[Na+].[Na+].[C:70]([O-:73])([OH:72])=O.[Na+].[CH2:75]1[CH2:79]OC[CH2:76]1>>[CH2:16]([C:58]([N:54]=[N+:55]=[N-:56])([C:57]([OH:60])=[O:59])[C:75]([CH3:79])([CH3:76])[C:70]([OH:73])=[O:72])[C:17]1[CH:18]=[CH:19][CH:20]=[CH:21][CH:22]=1Participants
Reactant 8




Product 1

Conditions
- Temperature
- 37.5 °C
Procedure
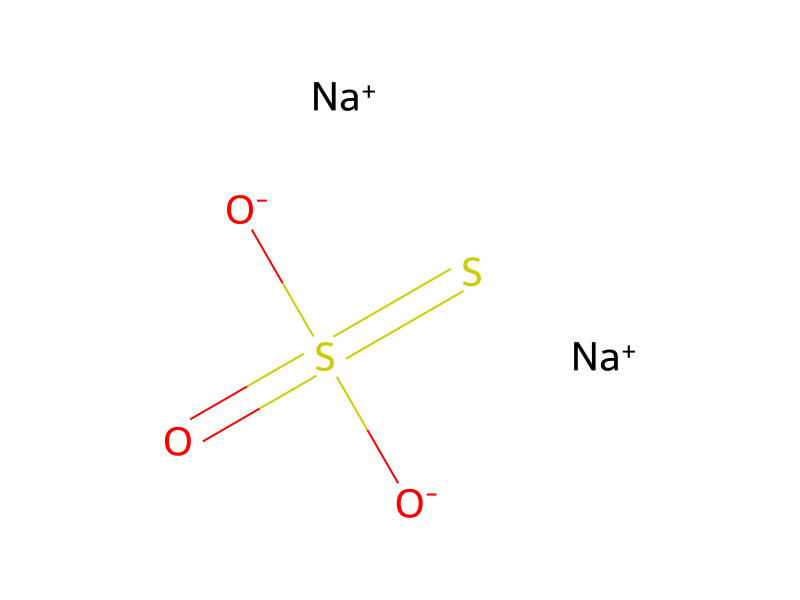
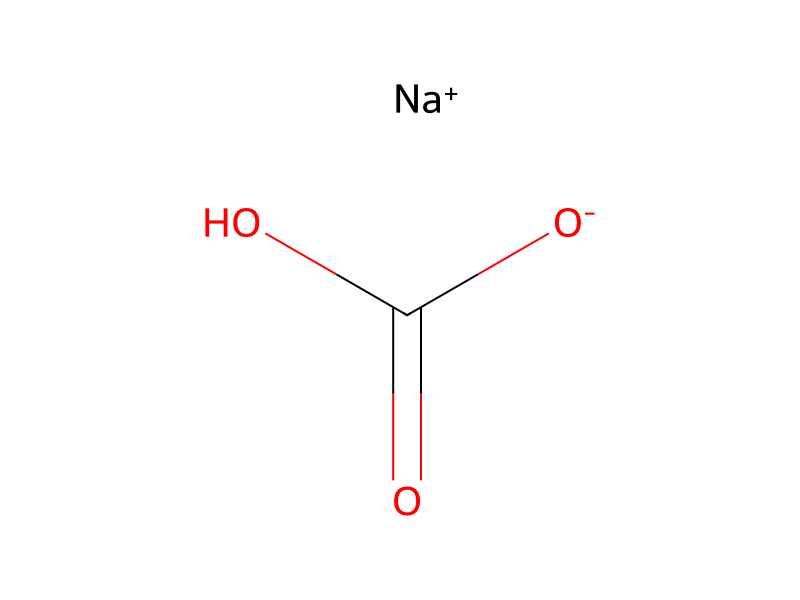
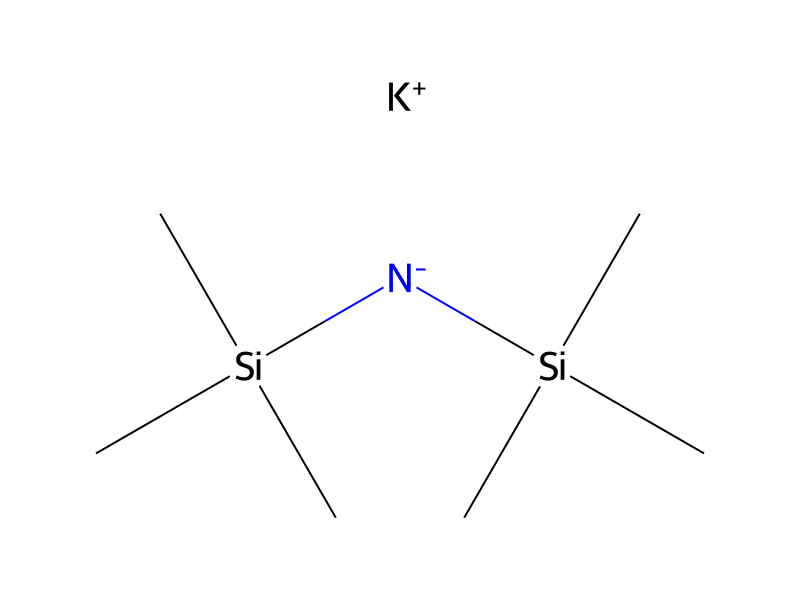
The oxazolidinone 34 (8.67 g, 24.9 mmol) was dissolved in THF (27 mL) and was added dropwise over 15 min to a solution of potassium bis(trimethylsilyl)amide (36.5 mL, 0.69 M in THF, 25.2 mmol) at −78° C. After 45 min at −78° C., 2,4,6-triisopropylbenzenesulfonyl azide (8.89 g, 28.7 mmol) in THF (15 mL) at −78° C. was added in one portion to the enolate. After 5 min, glacial acetic acid (4.6 equiv, 6.90 g, 6.56 mL, 0.12 mol) was added and the mixture was stirred at 35-40° C. for 90 min. Tetrahydrofuran was evaporated under reduced pressure and the residue was dissolved in a mixture of EtOAc and water. The organic phase was washed with saturated aqueous NaHCO3, brine, dried (MgSO4), filtered and concentrated. The residue was dissolved in 150 mL of hexane/ethyl acetate (2/1). After 16 h at 25° C., the white precipitate was removed by filtration and the filtrate was concentrated to give an oil which was filtered through a pad of silica and rinsed with hexane/ethyl acetate (5/1). This pale yellow oil (crude 35, 5.37 g, 55%) was dissolved in THF/water (260 mL, 3/1) and H2O2 (4 equiv, 30%, 6.24 mL, 55 mmol) was added at 0° C., followed by LiOHH2O (2.1 equiv, 1.22 g, 29 mmol). After 45 min, 10% aqueous Na2S2O3 (175 mL) and solid NaHCO3 (0.81 g) were added. Tetrahydrofuran was evaporated and the aqueous phase was extracted with chloroform (continuous liquid-liquid extraction, 24 h). The aqueous phase was then acidified with concentrated HCl at 0° C., and extracted with EtOAc (3 times). The combined EtOAc extracts were dried (MgSO4), filtered and concentrated. The residual oil was purified by flash chromatography on Merck silica gel eluting with ethyl acetate/acetic acid (400/1) to give compound 36 (0.82 g, 21%) as a colorless oil; [α]D22−74° (c 1.43, CHCl3); 1H-NMR (400 MHz, CDCl3) δ 7.39-7.32 (m, 5H), 5.20-5.12 (m, 2H), 4.48 (s, 1H), 1.34 (s, 3H), 1.29 (s, 3H); 13C-NMR (100.6 MHz, CDCl3) δ 174.74, 173.86, 135.42, 128.56, 128.35, 128.13, 67.86, 67.18, 45.98, 23.01, 20.29; FAB MS m/z: 278 (MH+); HRMS calcd for C13H16N3O4 (MH+) 278.1141, found: 278.1130; Anal (C13H15N3O4) C, H, N.
Workup
- waitAfter 5 min
- customTetrahydrofuran was evaporated under reduced pressure
- dissolutionthe residue was dissolved in a mixture of EtOAc and water
- washThe organic phase was washed with saturated aqueous NaHCO3, brine
- dry with materialdried (MgSO4)
- filtrationfiltered
- concentrationconcentrated
- dissolutionThe residue was dissolved in 150 mL of hexane/ethyl acetate (2/1)
- waitAfter 16 h at 25° C.
- customthe white precipitate was removed by filtration
- concentrationthe filtrate was concentrated
- customto give an oil which
- filtrationwas filtered through a pad of silica
- washrinsed with hexane/ethyl acetate (5/1)
- dissolutionThis pale yellow oil (crude 35, 5.37 g, 55%) was dissolved in THF/water (260 mL, 3/1)
- waitAfter 45 min
- customTetrahydrofuran was evaporated
- extractionthe aqueous phase was extracted with chloroform (continuous liquid-liquid extraction, 24 h)
- customwas then acidified with concentrated HCl at 0° C.
- extractionextracted with EtOAc (3 times)
- dry with materialThe combined EtOAc extracts were dried (MgSO4)
- filtrationfiltered
- concentrationconcentrated
- customThe residual oil was purified by flash chromatography on Merck silica gel eluting with ethyl acetate/acetic acid (400/1)
Sources & evidence
- ORD record
- ord-05fb29b926614ac082cbf7cc5b181a91
- Source dataset
- uspto-grants-2001_09
- Patent number
- US06291640B1
Reaction notes
Loading…