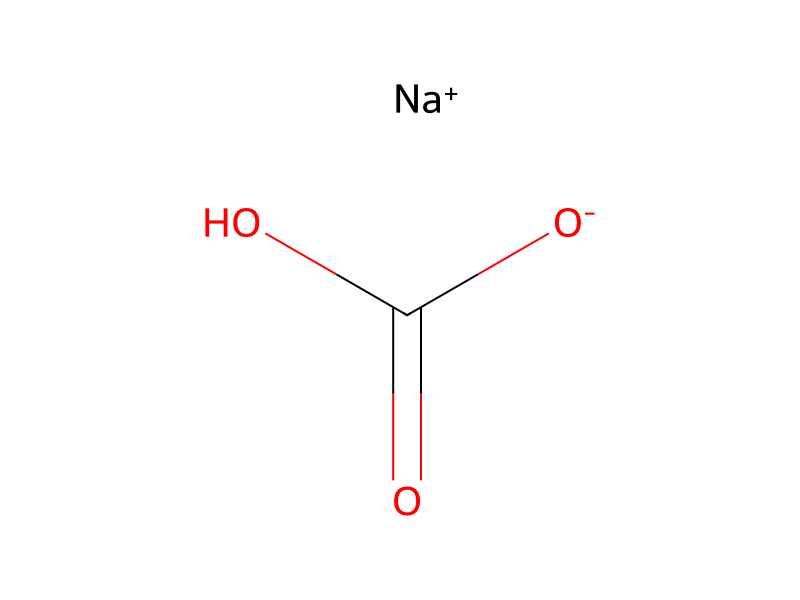
Sodium Bicarbonate → HCID 131733498, compound
Reaction equation

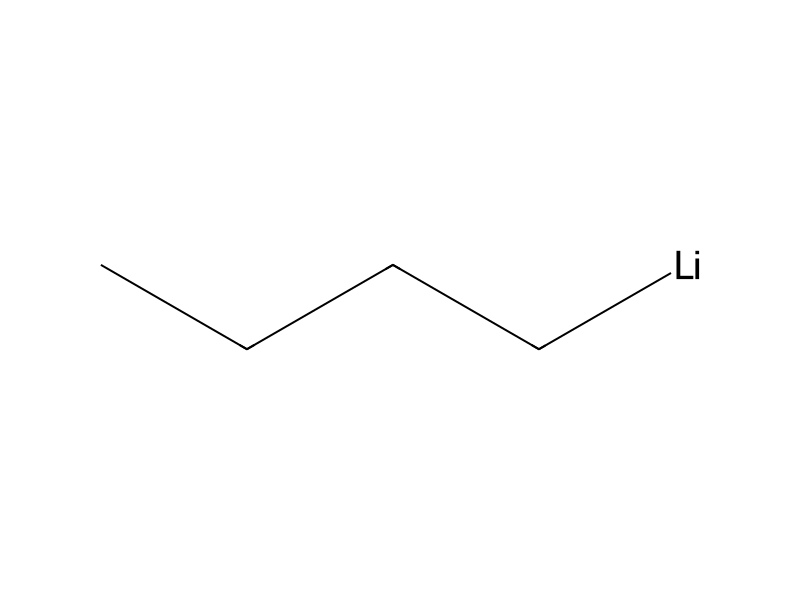
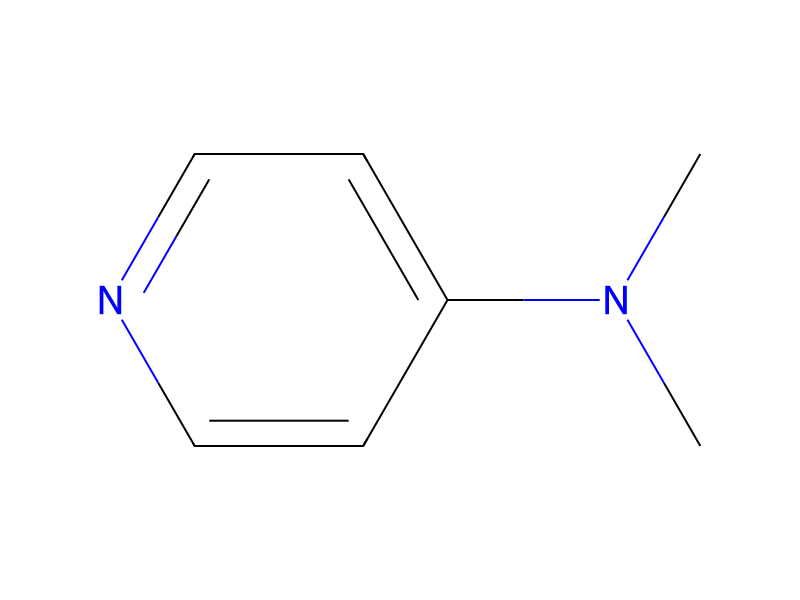
[C:1]([Si:5]([CH3:15])([CH3:14])[O:6][CH2:7][CH2:8][C:9]1[S:10][CH:11]=[CH:12][CH:13]=1)([CH3:4])([CH3:3])[CH3:2].C([Li])CCC.[Cl:21][C:22]1[CH:23]=[N+:24]([O-:42])[CH:25]=[C:26]([Cl:41])[C:27]=1[CH2:28][C@@H:29]([C:31]1[CH:36]=[CH:35][C:34]([O:37][CH3:38])=[C:33]([O:39][CH3:40])[CH:32]=1)[OH:30].Cl.CN(C)CCCN=C=NCC.[C:55](=O)(O)[O-:56].[Na+]>O1CCCC1.CN(C)C1C=CN=CC=1.ClCCl>[Cl:41][C:26]1[CH:25]=[N+:24]([O-:42])[CH:23]=[C:22]([Cl:21])[C:27]=1[CH2:28][C@H:29]([O:30][C:55]([C:11]1[S:10][C:9]([CH2:8][CH2:7][O:6][Si:5]([C:1]([CH3:2])([CH3:4])[CH3:3])([CH3:15])[CH3:14])=[CH:13][CH:12]=1)=[O:56])[C:31]1[CH:36]=[CH:35][C:34]([O:37][CH3:38])=[C:33]([O:39][CH3:40])[CH:32]=1Participants
Reactant 5


Product 1

Reagents, catalysts & solvents 3


Conditions
- Temperature
- 0 °C
Procedure
To a solution of tert-butyl-dimethyl-[2-(2-thienyl)ethoxy]silane synthesized as described in example 107 step 1 (485 mg, 2 mmol) in tetrahydrofuran (7.5 mL) at −78° C. was added n-butyl lithium (0.92 mL, 2.5 M in hexanes, 2.3 mmol) dropwise. The mixture was allowed to warm to 0° C. and allowed to stir at 0° C. for 1 h. The mixture was cooled to −78° C. and then gaseous CO2 was bubbled through the mixture for 30 minutes. The mixture was quenched by the addition of saturated aqueous ammonium chloride solution (15 mL), and allowed to warm to room temperature. The mixture was extracted with ethyl acetate (3×15 mL) and the combined organic fractions were passed through a hydrophobic fit and the solvent was removed by evaporation under reduced pressure. The residue was taken up in dichloromethane (4 mL) and (1S)-2-(3,5-dichloro-1-oxido-pyridin-1-ium-4-yl)-1-(3,4-dimethoxyphenyl)ethanol I-1/A (585 mg, 1.7 mmol), 4-dimethylaminopyridine (110 mg, 0.9 mmol) and N-(3-dimethylaminopropyl)-N′-ethylcarbodiimide hydrochloride (658 mg, 3.4 mmol) were added. The mixture was stirred at room temperature for 18 h and saturated aqueous sodium bicarbonate solution (10 mL) and dichloromethane (10 mL) were added. The layers were separated, and the organic phase was passed through a hydrophobic fit and the solvent was removed by evaporation under reduced pressure. The crude material was purified by silica gel column chromatography, eluting with 0-100% dichloromethane/10% methanol in dichloromethane, to afford the title compound (710 mg, 68%) as a brown oil.
Workup
- temperatureThe mixture was cooled to −78° C.
- customgaseous CO2 was bubbled through the mixture for 30 minutes
- customThe mixture was quenched by the addition of saturated aqueous ammonium chloride solution (15 mL)
- temperatureto warm to room temperature
- extractionThe mixture was extracted with ethyl acetate (3×15 mL)
- customthe solvent was removed by evaporation under reduced pressure
- stirringThe mixture was stirred at room temperature for 18 h
- customThe layers were separated
- customthe solvent was removed by evaporation under reduced pressure
- customThe crude material was purified by silica gel column chromatography
- washeluting with 0-100% dichloromethane/10% methanol in dichloromethane
Sources & evidence
- ORD record
- ord-20be4a3e7bb24819b2d5995feadaf864
- Source dataset
- uspto-grants-2015_09
- Patent number
- US09133185B2
Reaction notes
Loading…