Butyl iodide → HCID 19773340, compound
Reaction equation

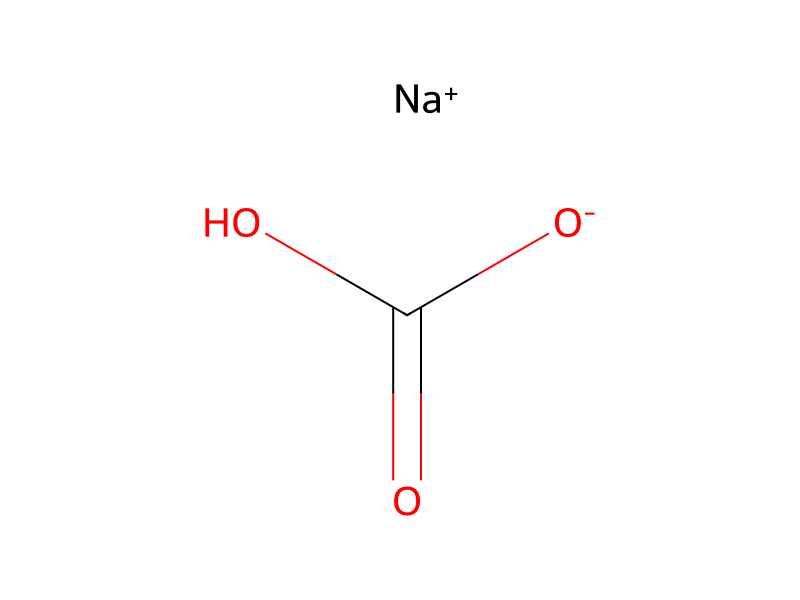
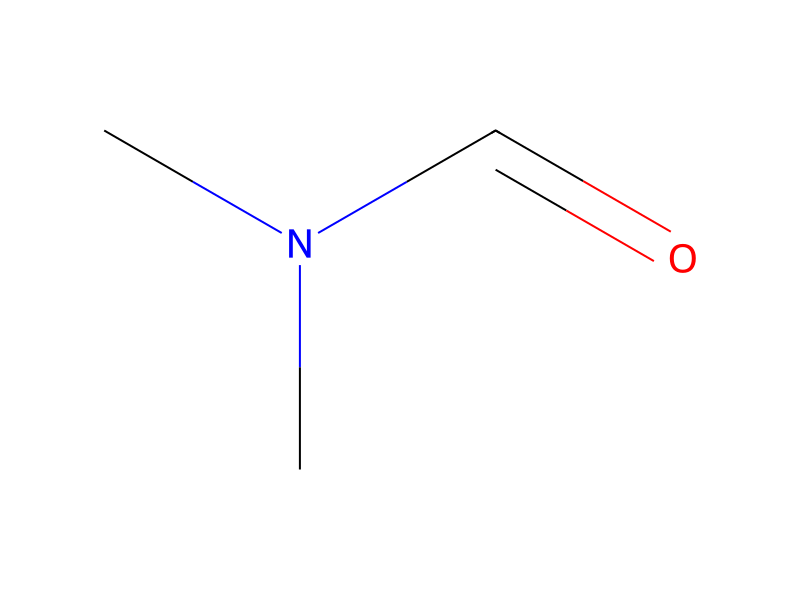
[ClH:1].[C:2]([NH:5][C:6]1[CH:15]=[CH:14][CH:13]=[C:12]2[C:7]=1[CH2:8][CH2:9][CH:10]([NH:16][CH2:17][CH2:18][CH3:19])[CH2:11]2)(=[O:4])[CH3:3].[CH2:20](I)[CH2:21][CH2:22][CH3:23].C(=O)([O-])O.[Na+]>CN(C)C=O.O1CCCC1>[ClH:1].[C:2]([NH:5][C:6]1[CH:15]=[CH:14][CH:13]=[C:12]2[C:7]=1[CH2:8][CH2:9][CH:10]([N:16]([CH2:20][CH2:21][CH2:22][CH3:23])[CH2:17][CH2:18][CH3:19])[CH2:11]2)(=[O:4])[CH3:3]Participants
Reactant 3


Product 1

Reagents, catalysts & solvents 2

Procedure
2.83 g (0.010 mol) of 5-acetylamino-2-n-propylamino-tetraline-hydrochloride (Example 4.1.1), 2.76 g (0.015 mol) of n-butyliodide and 1.68 g (0.020 mol) of sodium hydrogen carbonate are refluxed in a mixture of 20 ml of absolute dimethylformamide and 30 ml of absolute tetrahydrofuran for 48 hours with stirring (moment of complete reaction determined by thin layer chromatography). The reaction mixture is concentrated by rotary evaporation, finally at 95° C. under a complete water-jet vacuum. The residue is shaken with 50 ml of methylene chloride and 50 ml of water and the aqueous phase separated is extracted twice more with 15 ml of methylene chloride. The combined extracts are washed with 20 ml of water, dried with sodium sulphate and concentrated by rotary evaporation, finally at 90° C. under a complete water-jet vacuum. The residue is dissolved with 10 ml of ethanol and the solution is acidified with 4.0 ml of 2.5 N ethanolic hydrochloric acid (0.0I mol HCl). After the addition of diethylether until turbidity is just starting, the title compound crystallises. After standing overnight it is suction filtered at 5° C., washed with a mixture of ethanol and ether and dried at 80° C. until a constant weight is obtained. Yield 2.56 g (75.5% of theory), melting point 217° C. After recrystallisation from methanol/ether the melting point does not change.
Workup
- custom(moment of complete reaction determined by thin layer chromatography)
- concentrationThe reaction mixture is concentrated by rotary evaporation, finally at 95° C. under a complete water-jet vacuum
- stirringThe residue is shaken with 50 ml of methylene chloride and 50 ml of water
- customthe aqueous phase separated
- extractionis extracted twice more with 15 ml of methylene chloride
- washThe combined extracts are washed with 20 ml of water
- dry with materialdried with sodium sulphate
- concentrationconcentrated by rotary evaporation, finally at 90° C. under a complete water-jet vacuum
- dissolutionThe residue is dissolved with 10 ml of ethanol
- additionAfter the addition of diethylether until turbidity
- customthe title compound crystallises
- filtrationfiltered at 5° C.
- washwashed with a mixture of ethanol and ether
- customdried at 80° C. until a constant weight
- customis obtained
- customAfter recrystallisation from methanol/ether the melting point
Sources & evidence
- ORD record
- ord-b93eec9e567a4b57b45cd7bea262a9f3
- Source dataset
- uspto-grants-1993_03
- Patent number
- US05196454
Reaction notes
Loading…