Trifluoroacetic acid → 1,3-Dihydro-1-oxoisobenzofuran-5-carbonitrile
Reaction equation

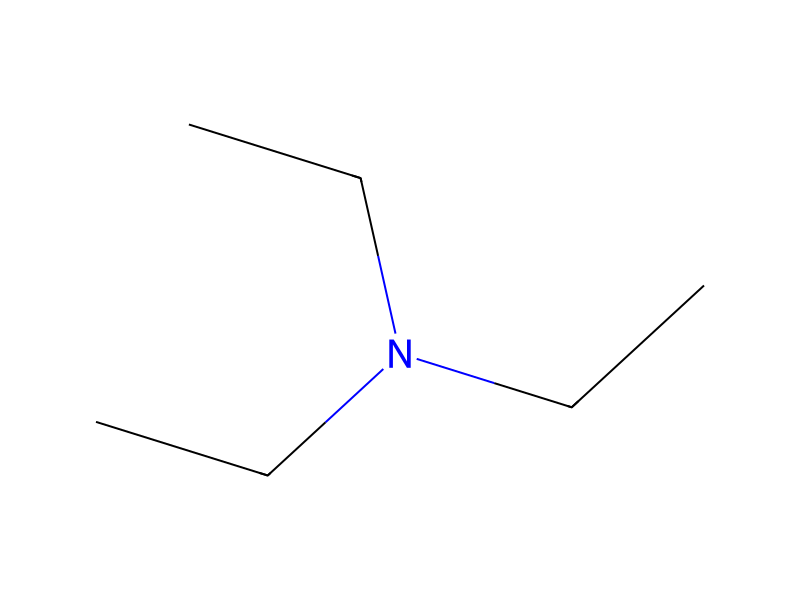
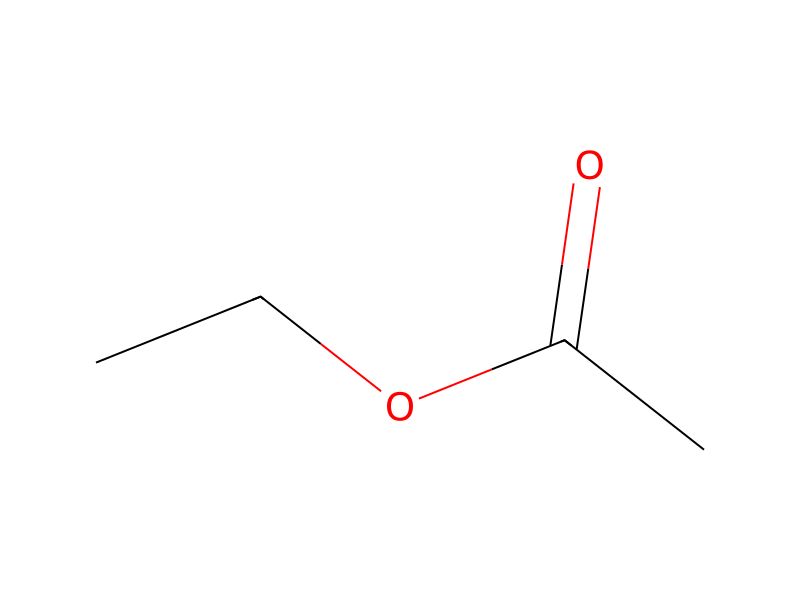
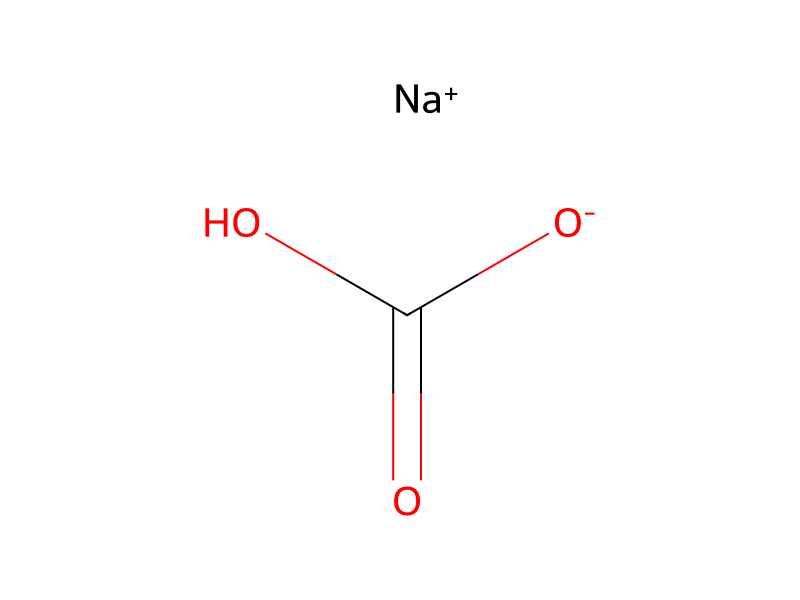
C([N:3]([CH2:6][CH3:7])CC)C.F[C:9](F)(F)[C:10](O)=O.[C:15](=O)([O-])O.[Na+].[C:20]([O:23][CH2:24][CH3:25])(=[O:22])[CH3:21]>O1CCCC1.ClCCl>[O:22]=[C:20]1[C:21]2[C:25](=[CH:15][C:7]([C:6]#[N:3])=[CH:9][CH:10]=2)[CH2:24][O:23]1Participants




Conditions
- Temperature
- 0 °C
Procedure
1-Oxo-1,3-dihydroisobenzofuran-5-carbaldehyde (2.01 g, 12.4 mmol) obtained from Example 4-(3) was suspended in tetrahydrofuran (50 ml), and after the suspension was cooled to 0° C., hydroxylamine hydrochloride (1.04 g, 14.9 mmol) in an aqueous solution of sodium hydroxide (1.0N; 14.8 ml, 14.8 mmol) was added thereto. The resulting mixture was stirred at room temperature for 1 hour, and then concentrated to one third of the volume under reduced pressure. To the concentrated solution was added water, and the product was extracted with ethyl acetate. The combined organic layers were washed with a saturated aqueous solution of sodium chloride, and the solvent was distilled off under reduced pressure to give crude 1-oxo-1,3-dihydroisobenzofuran-5-carbaldehyde oxime as a solid. The crude product was dissolved in tetrahydrofuran (50 ml), and the solution was cooled to 0° C., then triethylamine (3.04 g, 30 mmol) and anhydrous trifluoroacetic acid (3.13 g, 14.9 mmol) were added thereto. The resulting mixture was stirred at the same temperature for 30 minutes, and then warmed to room temperature followed by stirring for 30 minutes more. The reaction mixture was cooled again to 0° C., and a saturated aqueous solution of sodium hydrogen carbonate was added thereto. The reaction product was extracted with ethyl acetate, and the combined organic layers were washed with a saturated aqueous solution of sodium chloride. The extract was concentrated under reduced pressure to give a solid residue. The residue was subjected to chromatography on a silica gel (150 g) column (eluent; ethyl acetate:dichloromethane=0:1˜1:10) to afford the title compound (1.57 g, 79% yield) as a solid (mp. 200-201° C.).
Workup
- temperaturewarmed to room temperature
- stirringby stirring for 30 minutes more
- temperatureThe reaction mixture was cooled again to 0° C.
- extractionThe reaction product was extracted with ethyl acetate
- washthe combined organic layers were washed with a saturated aqueous solution of sodium chloride
- concentrationThe extract was concentrated under reduced pressure
- customto give a solid residue
Sources & evidence
- ORD record
- ord-f5f4c0ae299542039ab94a50ab85902a
- Source dataset
- uspto-grants-2007_06
- Patent number
- US07230023B2
Reaction notes
Loading…