Benzyl Bromide → Methyl 3-(benzyloxy)-2,2-dimethylpropanoate
Reaction equation

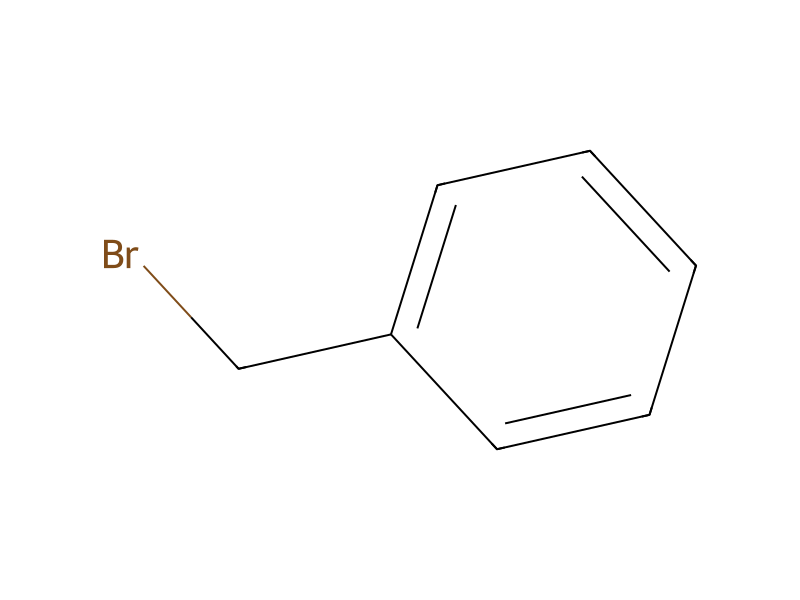
[OH:1][CH2:2][C:3]([CH3:9])([CH3:8])[C:4]([O:6][CH3:7])=[O:5].[H-].[Na+].[H][H].[CH2:14](Br)[C:15]1[CH:20]=[CH:19][CH:18]=[CH:17][CH:16]=1>CN(C)C=O.Cl.C(OC)(C)(C)C>[CH3:8][C:3]([CH3:9])([CH2:2][O:1][CH2:14][C:15]1[CH:20]=[CH:19][CH:18]=[CH:17][CH:16]=1)[C:4]([O:6][CH3:7])=[O:5]Participants
Reactant 4

Product 1

Reagents, catalysts & solvents 3
Conditions
- Temperature
- 60 °C
Procedure
Adapting procedures or variations thereof according to Eliel et al., J. Org. Chem., 1985, 50(15), 2707-2711; Roth et al., J. Agri. & Food. Chem. 1991, 39(3), 612-616; and Rega et al., Eur. J. Org. Chem. 2007, 6, 934-942, in an oven dried 500 mL round bottomed flask equipped with a magnetic stir bar and rubber septum, was added under a nitrogen atmosphere, 2.4 g of a 60 mass-% suspension of sodium hydride (NaH) in mineral oil (1.44 g of NaH, 60.0 mmol). The mineral oil was removed by washing with hexanes (Hxn) and the residue was dried under reduced pressure to yield a colorless solid (activated sodium hydride). Seven point nine (7.9) g (60 mmol) of methyl 3-hydroxy-2,2-dimethylpropanoate was added to a suspension of the activated sodium hydride in 200 mL of anhydrous N,N-dimethylformamide (DMF). The suspension was stirred until the hydrogen evolution ceased. Five-point-nine (5.9) mL (8.5 g, 50.0 mmol) of benzyl bromide (BnBr) was added and the mixture was stirred at ca. 60° C. for more than 10 hours. Upon completion and cooling, the reaction mixture was further diluted with one molar (1.0 M) hydrochloric acid (HCl) and methyl tert-butyl ether (MTBE) and the phases were separated. The aqueous phase was extracted two additional times with methyl tert-butyl ether (MTBE) and the combined organic extracts were washed with a saturated aqueous solution of sodium hydrogen carbonate (NaHCO3), water, and brine. The organic solution was dried over anhydrous magnesium sulfate (MgSO4), filtered, and the solvents were removed under reduced pressure using a rotary evaporator. The residue was further purified by silica gel column chromatography using a mixture of n-heptane (Hptn) and ethyl acetate (EtOAc) as eluent (EtOAc/Hptn=1:9) to provide 3.7 g (33% yield) of the title compound (26a) as a yellow oil. Rf=0.45 (EtOAc/Hptn=1:9). 1H NMR (400 MHz, CDCl3): δ=1.22 (s, 6H), 3.46 (s, 2H), 3.69 (s, 3H), 4.53 (s, 2H), 7.26-7.36 (m, 5H) ppm. The analytical data was consistent with the proposed structure.
Workup
- temperatureUpon completion and cooling
- customthe phases were separated
- extractionThe aqueous phase was extracted two additional times with methyl tert-butyl ether (MTBE)
- washthe combined organic extracts were washed with a saturated aqueous solution of sodium hydrogen carbonate (NaHCO3), water, and brine
- dry with materialThe organic solution was dried over anhydrous magnesium sulfate (MgSO4)
- filtrationfiltered
- customthe solvents were removed under reduced pressure
- customa rotary evaporator
- customThe residue was further purified by silica gel column chromatography
- additiona mixture of n-heptane (Hptn) and ethyl acetate (EtOAc) as eluent (EtOAc/Hptn=1:9)
Sources & evidence
- ORD record
- ord-bee3b82a768547da8a19315a464dce6d
- Source dataset
- uspto-grants-2012_05
- Patent number
- US08168617B2
Reaction notes
Loading…