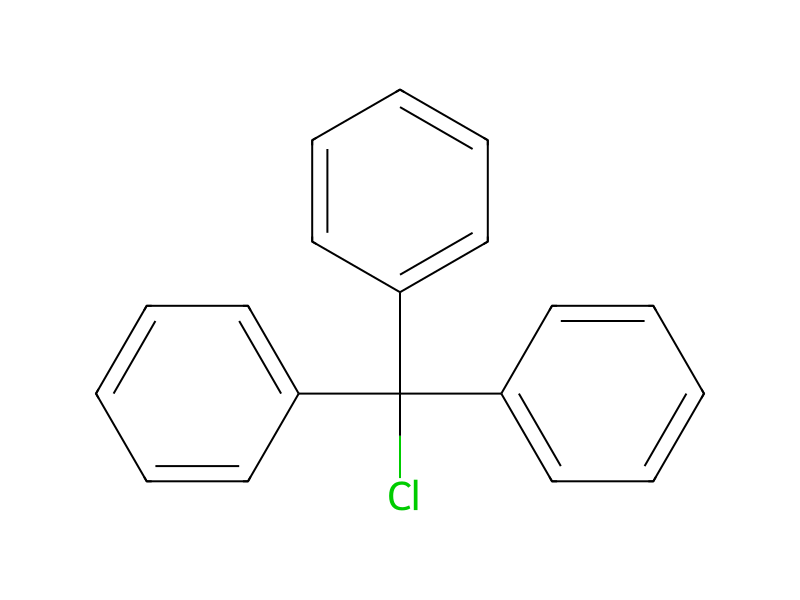
Trityl chloride → HCID 5274324, compound
Reaction equation

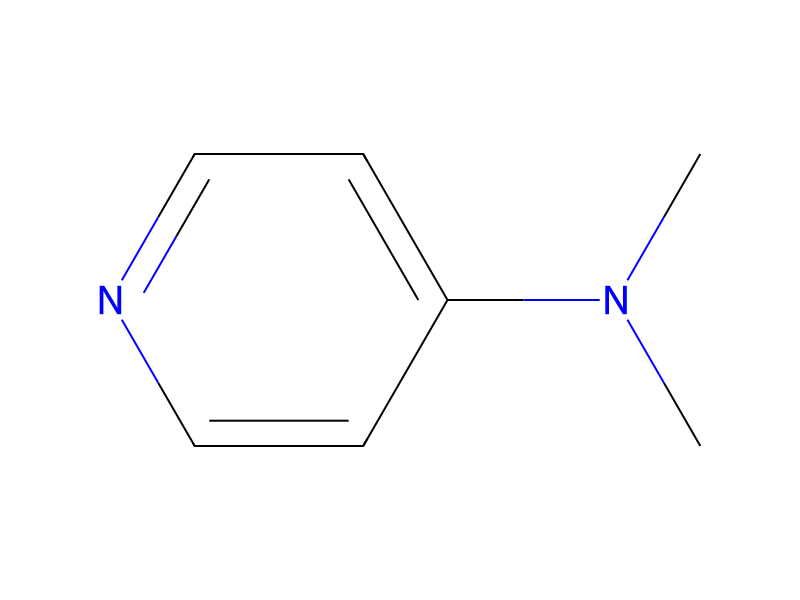
C(Cl)(C1C=CC=CC=1)(C1C=CC=CC=1)C1C=CC=CC=1.[OH:21][CH:22]([CH2:71][O:72]C(C1C=CC=CC=1)(C1C=CC=CC=1)C1C=CC=CC=1)[CH2:23][O:24][C:25]1[N:30]=[C:29]([NH:31]C(C2C=CC=CC=2)(C2C=CC=CC=2)C2C=CC=CC=2)[N:28]=[C:27]([NH:51]C(C2C=CC=CC=2)(C2C=CC=CC=2)C2C=CC=CC=2)[CH:26]=1.C1(C)C=CC(S(O[CH2:102][P:103](=[O:112])([O:108]C(C)C)[O:104]C(C)C)(=O)=O)=CC=1.[H-].[Na+]>CN(C)C1C=CN=CC=1.N1C=CC=CC=1.O1CCCC1.C(OCC)(=O)C.C(O)(=O)C.O>[NH2:31][C:29]1[N:28]=[C:27]([NH2:51])[CH:26]=[C:25]([O:24][CH2:23][CH:22]([O:21][CH2:102][P:103]([OH:112])([OH:108])=[O:104])[CH2:71][OH:72])[N:30]=1Participants
Reactant 4


Product 1

Reagents, catalysts & solvents 6


Conditions
- Temperature
- 50 °C
Procedure
The mixture of this compound (8.0 g, 40 mmol), trityl chloride (36.4 g, 131 mmol) and 4-dimethylaminopyridine (2 g) in pyridine (160 mL) was stirred 15 h at 50° C. and poured slowly under stirring to water (2 L). The slurry was stirred for 1 h, decanted, and, after stirring with fresh portion of water (2L) it was filtered and washed with water. The precipitate was taken up in chloroform (600 mL), dried with MgSO4 evaporated and codistilled with toluene (3×100 mL portions) in vacuo. The resulting gum was dissolved in minimum volume of ether and dropped, under vigorous stirring, to petroleum ether (1 L). The precipitate was filtered, washed with petroleum ether and dried. Yield, 33.4 g (91.5%) of 6-[2-hydroxy-3-(trityloxy)propoxy]-2,4-bis(tritylamino)pyrimidine, m.p. 138° C. For C64H54N4O2 (911.1) calculated 6.15% N; found 5.96% N. This trityl derivative (27.3 g, 30 mmol) and diisopropyl p-toluenesulfonyloxymethylphosphonate (15.75 g, 45 mmol) in tetrahydrofuran (300 mL) was treated with 60% NaH (5.4 g, 3.5 equivalents). The mixture was stirred at ambient temperature for 3 days and neutralized with acetic acid. The solvent was stripped off in vacuo, the residue dissolved in ethyl acetate (800 mL) and extracted with water (3×200 mL). The organic phase was evaporated to dryness and the residue refluxed in 80% aqueous acetic acid (300 mL) for 30 min, cooled and evaporated in vacuo. Water (300 mL) was added and the mixture extracted with ether (4×100 mL). The aqueous phase was freed of volatiles in vacuo and applied on a Dowex 50×8 column (200 mL) in acidic form. The column was washed with water till the drop of acidity and UV-absorbance and then eluted with 2.5% aqueous ammonia. The UV-absorbing ammonia eluate was collected, evaporated to dryness in vacuo, the residue codistilled with ethanol (3×100 mL) and dried over P2O5 in vacuo overnight. Acetonitrile (100 mL) and bromotrimethylsilane (30 mL) were added and the mixture was left to stand overnight under exclusion of moisture. After evaporation of the volatiles in vacuo, water (200 mL) was added to the residue, followed by conc. aqueous ammonia to an alkaline reaction. This solution was evaporated to dryness and the residue deionized on a Dowex 50×8 column (200 mL) under essentially the same conditions. The UV-absorbing ammonia eluate was taken down in vacuo, the residue was dissolved in minimum water and alkalized to pH 10 by ammonia. This solution was applied on a column (200 mL) Dowex 1×2 (acetate form) prewashed with water. The column was washed with water till the drop of UV-absorbance of the eluate and then with linear gradient of acetic acid (0-0,3 M, 1.5 1 each). The main UV-absorbing fraction was collected, evaporated in vacuo and the residue codistilled with water (2×100 mL). Crystallization of the residue from water gave 2,4-diamino-6-[(RS)-3-hydroxy-2-(phosphonomethoxy)propoxy]pyrimidine as monohydrate. Yield, 3.8 g (40.6%), white needles, m.p. 213° C. For C8H15N4O6P.H2O (312.2) calculated 30.78% C, 5.49% H, 17.94% N, 9.92% P; found 30.98% C, 5.52% H, 17.99% N, 9.82% P. Mass spectrum: 295.0 (MH+). 1H NMR (D2O+NaOD): 3.48 dd, 1H, J(P,CHb)=9.8, J(gem)=12.1 (P—CHb); 3.59 dd, 1H, J(P,CHa)=8.9, J(gem)=12.1 (P—CHa); 3.61 dd, 1H, J(3′b,2′)=6.8, J(gem)=12.9 (H-3′b); 3.70 m, 1H (H-2′); 3.71 dd, 1H, J(3′a,2′)=3.6, J(gem)=12.9 (H-3′a);4.11 dd, 1H, J(1′b,2′)=5.0, J(gem)=10.7 (H-1′b); 4.14 dd, 1H, J(1′a,2′)=4.6, J(gem)=10.7 (H-1′a); 5.36 s, 1H (H-5); 13C NMR (D2O+NaOD); 60.62 (C-3′); 65.58 (C-1′); 68.03 d, J(P,C)=149.4 (P—C); 76.63 (C-5); 79.89 d, J(P,C)=10.7 (C-2′); 162.63 and 166.54 (C-2, C-4); 170.78 (C-6).
Workup
- additionpoured slowly
- stirringThe slurry was stirred for 1 h
- customdecanted
- stirringafter stirring with fresh portion of water (2L) it
- filtrationwas filtered
- washwashed with water
- dry with materialdried with MgSO4
- customevaporated
- dissolutionThe resulting gum was dissolved in minimum volume of ether
- additiondropped, under vigorous stirring, to petroleum ether (1 L)
- filtrationThe precipitate was filtered
- washwashed with petroleum ether
- customdried
- customYield
- stirringThe mixture was stirred at ambient temperature for 3 days
- extractionextracted with water (3×200 mL)
- customThe organic phase was evaporated to dryness
- temperaturethe residue refluxed in 80% aqueous acetic acid (300 mL) for 30 min
- temperaturecooled
- customevaporated in vacuo
- additionWater (300 mL) was added
- extractionthe mixture extracted with ether (4×100 mL)
- washThe column was washed with water till the drop of acidity and UV-absorbance
- washeluted with 2.5% aqueous ammonia
- customThe UV-absorbing ammonia eluate was collected
- customevaporated to dryness in vacuo
- dry with materialdried over P2O5 in vacuo overnight
- additionAcetonitrile (100 mL) and bromotrimethylsilane (30 mL) were added
- waitthe mixture was left
- waitto stand overnight under exclusion of moisture
- customAfter evaporation of the volatiles in vacuo, water (200 mL)
- additionwas added to the residue
- customfollowed by conc. aqueous ammonia to an alkaline reaction
- customThis solution was evaporated to dryness
- dissolutionthe residue was dissolved in minimum water
- washThe column was washed with water till the drop of UV-absorbance of the eluate
- customThe main UV-absorbing fraction was collected
- customevaporated in vacuo
- customCrystallization of the residue from water
Sources & evidence
- ORD record
- ord-9fd782d547944804a32a74acc0c1ab80
- Source dataset
- uspto-grants-2004_11
- Patent number
- US06818633B2
Reaction notes
Loading…